

 menu


- 1. Introduction
- 2. EMBL Hamburg origins and beamline deployment
- 3. Architecture: a web interface coupled to crystallographic decision makers
- 4. Phasing strategies supported by Auto-Rickshaw
- 5. SHELX as the main experimental phasing engine
- 6. Auto-Rickshaw in a wider automation landscape
- 7. Auto-Rickshaw at the beamline
- 8. Scientific use and community impact
- 9. Evolution: MRSAD, RIP, S-SAD and multi-dataset workflows
- 10. Current status and outlook of the Auto-Rickshaw web service
- 11. From Auto-Rickshaw to ASWEBRICK: secure and reproducible deployment
- 12. Auto-Rickshaw in the current landscape
- 13. Lessons from Auto-Rickshaw's design philosophy
- 14. Conclusions
- References
A Tribute to George Sheldrick
 | STRUCTURAL BIOLOGY |
 access
A short history of Auto-Rickshaw

aANSTO, Australian Synchrotron, 800 Blackburn Road, Clayton, Victoria 3168, Australia, bDepartment of Biochemistry and Molecular Biology, Monash University, Victoria 3800, Australia, cEuropean Molecular Biology Laboratory, Hamburg Unit, Hamburg, Germany, dEuropean Molecular Biology Laboratory, Heidelberg and Hamburg Units, Germany, and eMacromolecular Crystallography, Helmholtz-Zentrum Berlin, Albert-Einstein-Strasse 15, 12489 Berlin, Germany
*Correspondence e-mail: [email protected]
This article is part of the collection A Tribute to George Sheldrick: the Legacy of a Crystallographic Computing Pioneer.
Auto-Rickshaw was developed in the early 2000s at EMBL Hamburg as an automated macromolecular structure-determination platform for crystallography and structural biology. Its initial motivation was to provide beamline users with rapid feedback on whether diffraction data were sufficient for structure solution. The platform was not merely a wrapper around existing crystallographic programs; it encoded the expert decisions needed to select phasing, density-modification, model-building and refinement protocols according to the information available from the experiment. Since its first publication in 2005, Auto-Rickshaw has supported a broad range of experimental and model-based strategies, including SAD, MAD, SIR, SIRAS, RIP, molecular replacement and hybrid approaches such as MRSAD. A central feature was the integration of the SHELX programs, particularly SHELXC, SHELXD and SHELXE, whose speed, robustness and command-line design made them especially suitable for automated pipelines. This review gives a brief historical account of Auto-Rickshaw, the beamline context in which it emerged, the design principles that shaped it and the role of SHELX programs in its phasing architecture. It also considers the broader legacy of Auto-Rickshaw as a bridge between crystallographic expertise and automated, reproducible structure solution. Finally, the review places Auto-Rickshaw in the current landscape of synchrotron technologies, high-throughput structural biology, containerized deployment, predicted structural models and AI-assisted decision making, where reliable automation remains essential.
Keywords: Auto-Rickshaw; experimental phasing; molecular replacement; MRSAD; SHELX; macromolecular crystallography; structural biology; SHELXC; SHELXD; SHELXE; synchrotron beamlines; automation.
1. Introduction
For many decades, the crystallographic phase problem constituted the central obstacle to structure determination of biological macromolecules by X-ray crystallography. Diffraction experiments measure reflection intensities, from which structure-factor amplitudes can be derived, but the corresponding phases are not measured directly. Solving a structure therefore requires additional information: anomalous differences, isomorphous differences, radiation-induced changes, prior structural models, noncrystallographic symmetry, density-modification constraints, or some combination of these. The conceptual basis of these approaches is well established, but their practical use has historically required experience, judgement and persistence.
Before the widespread adoption of automated structure-solution pipelines, a crystallographer solving a new macromolecular structure had to make a long sequence of interconnected decisions. Which phasing strategy was appropriate? Was the anomalous signal significant? What resolution cutoff should be used for substructure solution? How many heavy-atom or anomalous-scatterer sites should be expected? Was the substructure hand correct? Was the electron-density map interpretable? Should molecular-replacement phases be combined with anomalous data? When should density modification, model building or refinement be attempted? Each of these choices could determine whether a structure was solved rapidly, solved only after many iterations, or missed entirely.
The development of Auto-Rickshaw greatly benefitted from the DESY/EMBL Hamburg macromolecular crystallography (MX) beamline environment, where software automation, user support and experimental decision making were closely interconnected. At the time, EMBL Hamburg operated five MX beamlines at the DORIS/DORIS III storage ring, a second-generation synchrotron source: BW7A, BW7B, X11, X12 and X13. These beamlines provided complementary experimental capabilities: BW7A and later X12 were tuneable beamlines used for anomalous diffraction experiments, whereas BW7B, X11 and X13 primarily supported fixed-wavelength data collection and rapid high-resolution or single-wavelength anomalous diffraction (SAD) experiments. Thus, Auto-Rickshaw was developed in an environment that included both wavelength-tuneable experimental phasing beamlines and fixed-wavelength stations (Pohl et al., 2001 , 2004; Hermes, 2004).
Moreover, many researchers from the group of George Sheldrick at the University of Göttingen regularly visited EMBL Hamburg for crystallographic experiments and this strongly contributed to the development of methodological innovations.
Data collection at the time was also substantially more beamtime-consuming than in the present era of high-brilliance sources equipped with fast pixel-array detectors. Depending on the crystal, detector, strategy and phasing experiment, a single dataset could require tens of minutes to hours. In that setting, rapid computational validation was not merely a convenience: it could affect the experiment while beamtime was still available, for example by showing that sufficient phasing information had already been collected, or that additional data, a different wavelength or another crystal was required.
Auto-Rickshaw was designed as an automated crystal-structure determination platform that could guide diffraction data through alternative phasing and model-building routes. The name itself has a practical origin. In early internal descriptions, the system was referred to more formally as the EMBL Hamburg automated crystal-structure determination platform. During review of the original manuscript, however, the authors were encouraged to give the platform a distinctive name. The term Auto-Rickshaw had already been used colloquially within the group because it captured the intended behaviour of the software. In South Asia, an auto-rickshaw, also known in some regions as a tuk-tuk, is a motorized successor to a manually driven rickshaw. By analogy, the program was intended to automate many of the crystallographic decisions that had previously been made manually: choosing routes, responding to obstacles and moving efficiently through a complex structure-solution landscape.
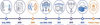
The goal was not to replace crystallographic expertise, but to make expert-like workflows available to non-expert users in a reliable, reproducible and accessible form. This review summarizes the circumstances of its development, the principles behind its architecture, the evolution of its phasing protocols and the role of SHELX programs. It follows Auto-Rickshaw from a public web service to beamline-integrated, secured and containerized implementations, as summarized in Fig. 1, and places it alongside contemporary systems such as SOLVE/RESOLVE, HKL-3000, APRV, autoSHARP, CRANK and ELVES.
|
Figure 1
Historical timeline of Auto-Rickshaw. The timeline illustrates key milestones in the evolution of Auto-Rickshaw from its EMBL Hamburg/DESY beamline origins in the early 2000s through modern secure and containerized deployment. Major developments include the 2005 initial release, the introduction of MRSAD in 2009, relocation to the Australian Synchrotron in 2011 and containerized deployment by 2021. A detailed description of each phase is provided in the text. |
2. EMBL Hamburg origins and beamline deployment
Auto-Rickshaw was conceived and developed at the EMBL Hamburg Outstation on the DESY campus in Hamburg, a research environment that also served a large international beamline user community. The platform therefore emerged from practical synchrotron needs as much as from software development itself.
At the time, the EMBL Hamburg MX beamlines at DESY formed one of Europe's major synchrotron MX facilities. BioSync/PDB beamline statistics for deposition years 1995–2009 list more than 2000 publicly available macromolecular PDB depositions under the EMBL/DESY entry, corresponding to approximately one-sixth of all European depositions. EMBL/DESY was the second-largest European synchrotron structure provider, after the third-generation source ESRF (BioSync, 2026).
During beamtime visits, users often had to decide quickly whether to continue with the current data, collect more data, change strategy or try another crystal. Auto-Rickshaw was designed to support those decisions by turning processed diffraction data into rapid phasing attempts, maps, partial models and diagnostic information.
Auto-Rickshaw arose from this practical beamline setting. The developers recognized that many parts of expert crystallographic decision making could be encoded as a structured workflow. In its initial form, the pipeline focused primarily on automated experimental phasing from anomalous and derivative data, including SAD, MAD and SIRAS strategies, together with the associated steps of data preparation, substructure determination, phase improvement, density modification, model building and refinement. The system therefore needed to accept user data, analyse the available phasing information, select appropriate software tools, monitor intermediate results and redirect the calculation if necessary (Panjikar et al., 2005).
The development also belonged to the broader European high-throughput structural biology environment of the early 2000s. Support from the EC-funded BIOXHIT and SPINE projects connected automated structure solution, synchrotron operation and structural genomics (Panjikar et al., 2005).
The subsequent 2009 introduction of MRSAD, the combination of molecular replacement with SAD phasing, became one of the most important extensions of Auto-Rickshaw and is discussed in Section 9 below (Panjikar et al., 2009).
After the relocation of the principal developer (SP) to the Australian Synchrotron in June 2011, the EMBL Hamburg public web service continued to be maintained and further beamline-focused development proceeded at the Australian Synchrotron, where Auto-Rickshaw could be used for rapid data evaluation and user support (Australian Synchrotron, 2011; Panjikar, 2021).
As detector readout speed, beamline automation and sample throughput increased, the platform also evolved towards multiple-dataset evaluation. Subsequent developments introduced `multiscale' and `multidata' modes, in which datasets could be selected, clustered, scaled or merged according to unit-cell similarity, space group, isomorphism, radiation damage and anomalous signal before being passed to SAD, SIRAS or MAD phasing workflows (Panjikar et al., 2015, 2017). This extended the original single-job validation concept towards beamline-scale data selection and automated evaluation of many related datasets.
The Australian Synchrotron deployment also reflected the wider move towards high-performance and containerized scientific computing, including the 2021 installation on the ASCI cluster using Docker and Kubernetes and the later ASWEBRICK (Australian Synchrotron WEB-based RICKshaw) development described below. These technologies should be understood as later deployment and orchestration mechanisms rather than as components of the original Auto-Rickshaw decision-making methodology.
3. Architecture: a web interface coupled to crystallographic decision makers
The original Auto-Rickshaw architecture combined a web-based graphical interface with a backend web service that controlled crystallographic programs. Users supplied processed diffraction data, sequence information and experimental details without installing the underlying software locally; the backend connected data preparation, substructure solution, phasing, density modification, model building and refinement.
The system evaluated intermediate outputs and used them to choose subsequent steps. In experimental phasing, the strength of the anomalous or isomorphous signal, substructure quality, hand discrimination, density-modification success and map interpretability could all influence the path forward.
In practical terms, Auto-Rickshaw selected among protocols, tested hands or resolution limits, chose downstream programs, stopped weak routes and reported diagnostic evidence. It did not learn during an individual run; its rules evolved through successive releases as new protocols, diagnostics and crystallographic experience were incorporated.
A clear example of this decision logic was the selection of heavy-atom parameter refinement and phase-calculation programs according to quantitative indicators such as the correlation coefficient for the weak signal, solvent content and data resolution. In the original implementation, these criteria were used to choose between alternative downstream programs, including BP3 (Pannu & Read, 2004), SHARP (de La Fortelle & Bricogne, 1997), MLPHARE (Winn et al., 2011) and SHELXE, or to halt the calculation when the signal or heavy-atom model was too weak to justify further automated processing (Panjikar et al., 2005). The precise numerical boundaries between these decision regions were not intended to be immutable; rather, they represented an evolving empirical description of expert crystallographic judgement. This feature is central to the historical importance of Auto-Rickshaw: it encoded not only software execution, but also the conditional choices that crystallographers make when assessing whether experimental phasing information is strong enough to proceed.
Auto-Rickshaw was made available in two flavours. The beamline, or fast, version was optimized for rapid assessment of diffraction experiments, whereas the advanced version was designed for more complete structure determination and required additional information, including sequence data.
The backend integrated a wide range of crystallographic programs. Data-processing and scaling tools were linked to phasing programs, molecular-replacement programs, density-modification tools, automated model-building programs and refinement packages. The SHELX programs occupied a particularly central position in experimental phasing. Molecular replacement was supported through established tools such as MOLREP (Vagin & Teplyakov, 1997) and Phaser (McCoy et al., 2007), while density modification, model building and refinement could involve programs such as DM (Winn et al., 2011), RESOLVE (Terwilliger, 2000), ARP/wARP (Lamzin & Wilson, 1993; Langer et al., 2008), Buccaneer (Cowtan, 2006), Coot (Emsley et al., 2010), REFMAC (Murshudov et al., 1997) and phenix.refine (Adams et al., 2010).
4. Phasing strategies supported by Auto-Rickshaw
One of the defining features of Auto-Rickshaw was the breadth of phasing strategies that it supported. Rather than focusing on a single method, the platform was designed to select among several experimental and model-based routes according to the information available from the diffraction experiment.
Single-wavelength anomalous diffraction (SAD) became one of the most important protocols. In SAD experiments, anomalous differences from selenium, sulfur, halides or other anomalous scatterers provide the information needed to locate a substructure and estimate phases. Auto-Rickshaw automated the complete sequence: data preparation and signal analysis, substructure solution, phase improvement, model building and refinement.
Multi-wavelength anomalous diffraction (MAD) was also supported, including experiments with two, three or four wavelengths. MAD experiments were historically central to synchrotron-based macromolecular crystallography. An early landmark was the de novo structure determination of a basic blue copper protein from cucumber seedlings using multiple-wavelength anomalous diffraction (Guss et al., 1988). This and subsequent MAD applications demonstrated the value of tuneable synchrotron beamlines, where data could be collected at peak, inflection and remote wavelengths around an absorption edge. Auto-Rickshaw provided a way to manage the additional complexity of multi-wavelength phasing, including wavelength-specific data preparation and substructure determination.
Isomorphous replacement methods, including single isomorphous replacement (SIR) and single isomorphous replacement with anomalous scattering (SIRAS), formed another component of the platform. These methods exploit differences between native and derivative crystals, sometimes combined with anomalous differences. They are sensitive to non-isomorphism and derivative quality, making them well suited to automated diagnostic evaluation.
Molecular replacement (MR) was included because, as the Protein Data Bank expanded, MR became an increasingly dominant route for macromolecular structure solution. This trend was already clear in the late 2000s: the BALBES molecular-replacement pipeline was developed to exploit the growing PDB as a search-model repository, and testing of that pipeline indicated that around 75% of all structures could be solved automatically without user intervention (Long et al., 2008). A later discussion of MOLREP noted that about two-thirds of recently deposited X-ray structures in the PDB had been determined by MR (Vagin & Teplyakov, 2010). Auto-Rickshaw therefore included MR programs to obtain initial model phases and then connected these phases to refinement, density modification and automated model building.
Hybrid and specialist routes were also supported. These included molecular replacement combined with SAD phasing (MRSAD), radiation-induced phasing (RIP), molecular-replacement-assisted RIP (MRRIP), sulfur SAD (S-SAD) and later multiple-dataset workflows. These extensions are discussed in more detail in Section 9.
Table 1 summarizes the phasing methods supported by Auto-Rickshaw and the principal SHELX programs involved in each route.
|
|||||||||||||||||||||||||||||
5. SHELX as the main experimental phasing engine
The SHELX suite, developed by George M. Sheldrick, became important to Auto-Rickshaw because its programs combined algorithmic power with the practical qualities needed for automation: speed, robustness, simple input and output, and command-line operation. The triad SHELXC, SHELXD and SHELXE was especially important for experimental phasing (Sheldrick, 2008, 2010; Usón & Sheldrick, 2018, 2024).
SHELXC provided data preparation and analysis. It estimated anomalous or marker-atom structure-factor amplitudes, generated normalized structure factors and produced input for subsequent SHELX programs. Equally importantly, it provided statistical diagnostics that could be used by Auto-Rickshaw decision makers. Measures of anomalous correlation and signal strength helped determine whether a dataset was likely to support substructure solution and to what resolution the signal remained useful.
SHELXD solved the heavy-atom or anomalous-scatterer substructure using dual-space direct methods (Schneider & Sheldrick, 2002). In the context of automated pipelines, SHELXD was valuable not only because it was powerful, but because it was fast enough to be run repeatedly under different assumptions. Auto-Rickshaw could explore alternative resolution cutoffs, expected site numbers or data treatments, increasing the chance of finding a correct substructure without requiring a user to perform trial-and-error searches manually.
SHELXE provided phase improvement, density modification and, crucially, automated chain tracing (Sheldrick, 2010; Usón & Sheldrick, 2018, 2024). The ability of SHELXE to distinguish the correct hand of a substructure and to produce interpretable density or partial polyalanine models made it an excellent diagnostic and productive step in an automated pipeline. In many cases, the transition from an abstract substructure solution to a map that a crystallographer could recognize as protein-like occurred through SHELXE.
More recent developments in SHELXE further emphasize why it remains relevant to the Auto-Rickshaw philosophy. SHELXE can operate with phases derived from experimental data, from partial models or maps, or from combinations of these sources, including workflows such as MRSAD. It can also read external phase values, phase weights or phase-probability distributions encoded as Hendrickson–Lattman coefficients. These capabilities make SHELXE well suited to an adaptive pipeline in which phase information may arise from different sources and at different stages of the calculation. The recent extension of tracing into side chains and the explicit treatment of model bias from predicted models also connect SHELXE to current developments in AlphaFold-assisted molecular replacement and model completion (Usón & Sheldrick, 2024).
Within Auto-Rickshaw, the use of SHELXE was therefore not simply automatic in every case. It was selected as part of a broader decision framework in which signal strength, solvent content, resolution and heavy-atom model quality influenced whether SHELXE, MLPHARE, BP3 or SHARP was used for subsequent phase calculation or phase improvement (Panjikar et al., 2005). This historical detail is important because it shows that the central role of SHELX in Auto-Rickshaw was embedded within a wider adaptive phasing strategy.
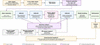
The overall experimental phasing workflow is illustrated in Fig. 2. In this workflow, SHELX programs served as dependable engines within a larger decision-making framework: Auto-Rickshaw provided the route planning, while SHELX provided much of the experimental phasing horsepower.
|
Figure 2
Experimental phasing workflow in Auto-Rickshaw. User input and diffraction data are passed through the web-server/job-control layer to protocol selection, which determines the appropriate data-reduction and phasing route. The central SHELX-based path uses SHELXC for data preparation and signal analysis, SHELXD for anomalous or heavy-atom substructure solution and SHELXE for hand evaluation, phase improvement, density modification and autotracing. Decision makers monitor intermediate results, test alternative resolution cutoffs and hands, and reroute unsuccessful paths. When a search model is provided, phased molecular replacement is carried out using MOLREP or Phaser. The resulting MR phases or partial model can assist the selected experimental phasing protocol before phase improvement, density modification and autotracing in SHELXE. Final maps, models and diagnostic reports inform subsequent experimental decisions. |
6. Auto-Rickshaw in a wider automation landscape
Auto-Rickshaw did not emerge in isolation. Its development formed part of a broader movement in macromolecular crystallography towards the automation of data processing, experimental phasing, model building and refinement. Contemporary and near-contemporary systems included autoSHARP, ACrS, HKL-3000, ELVES, APRV, CRANK and SOLVE/RESOLVE, each providing a different degree of automation and addressing a different part of the crystallographic workflow (Vonrhein et al., 2007; Brunzelle et al., 2003; Minor et al., 2006; Holton & Alber, 2004; Kroemer et al., 2004; Ness et al., 2004; Terwilliger, 2000, 2004).
Table 2 summarizes these systems and their relation to Auto-Rickshaw. The distinguishing emphasis of Auto-Rickshaw was web-accessible, decision-driven routing across multiple programs and phasing strategies, with feedback fast enough to support beamline decisions.
|
||||||||||||||||||||||||||||||||
7. Auto-Rickshaw at the beamline
At a synchrotron beamline, time is limited, data quality varies and decisions often need to be made while the experiment is still in progress. The beamline version of Auto-Rickshaw connected processed data to automated phasing and preliminary model building, helping users decide whether to continue collecting data, continue measurements at another wavelength of the radiation, collect a derivative, increase multiplicity or move to another sample.
Two representative beamline examples illustrate this point. In the first case, reported as real case R7 in the original Auto-Rickshaw paper (Panjikar et al., 2005), a selenium-labelled protein diffracting to 2.55 Å resolution with high solvent content was solved sufficiently rapidly to influence the ongoing experiment. This case corresponds to the aryl-alcohol oxidase structure from Pleurotus eryngii, later deposited as PDB entry 3fim (Fernández et al., 2009). After substructure solution, the Auto-Rickshaw decision makers selected SHELXE for phase calculation and continued with automated model building in ARP/wARP. The Beamline Version produced an interpretable map and a partial α-helical model with 363 of 593 residues built while data collection was still in progress; at that point, the user halted collection of the second wavelength of a planned MAD experiment because the space-group ambiguity had already been resolved and the solvent content correctly estimated. The Advanced Version subsequently built 87% of the model. The difference between the 593 residues described for the R7 case and the 566 residues reported in the deposited structure reflects poorly ordered regions in the final deposited coordinate model. This example shows how Auto-Rickshaw could demonstrate that fewer data than originally planned were sufficient for structure solution and the valuable beamtime saved could be used for other experiments.
A complementary beamline scenario illustrates the opposite outcome. In a second heavy-atom case, initial SAD, two-wavelength MAD and three-wavelength MAD attempts did not produce a solution. A successful structure-solution route was obtained only after higher redundancy anomalous data had been collected. Although this example was reported as an operational beamline/teaching case rather than as a separate structure report, it captures an important use of automated feedback: Auto-Rickshaw could indicate that the current data were insufficient and that additional data, rather than continuation of the original strategy, were required (Panjikar, 2010).
These representative scenarios are summarized in Table 3. This feedback addressed practical questions faced during an experiment and also recorded the programs used, the phasing route selected and key intermediate decisions. At the end of a run, Auto-Rickshaw listed the relevant publications to cite for the software and protocols used, helping users prepare the structure-solution description for publication.
|
|||||||||||||||||
As synchrotron MX moved towards automated sample handling, remote access and high-throughput data collection, this role became more valuable. Structural genomics programmes, fragment-screening projects and mail-in crystallography all created pressure for reproducible, scalable and minimally supervised workflows. Auto-Rickshaw lowered the barrier to experimental phasing, but its output still required crystallographic judgement: models, maps, substructures and failed runs all had to be interpreted.
8. Scientific use and community impact
The scientific impact of Auto-Rickshaw is reflected not only in its methodological role, but also in its sustained use by the macromolecular crystallography community. By providing a web-accessible and decision-driven structure-solution platform, Auto-Rickshaw lowered the barrier to experimental phasing and enabled users to test multiple phasing routes without having to install, configure and manually coordinate a large number of crystallographic programs.
Cumulative server and citation statistics for the period from 2005 to August 2023 provide a useful measure of this community use and are summarized in Table 4. The data demonstrate that Auto-Rickshaw became a practical resource for both routine and challenging structure-solution problems across a large international user base. The distribution of submitted jobs illustrates the practical breadth of the platform: SAD and MR accounted for the largest fractions of use, while MRSAD, MAD, S-SAD and RIP protocols provided important additional routes for more specialized or difficult cases.
|
||||||||||||||||||||||
These statistics should therefore be interpreted as evidence of broad community adoption rather than as a direct count of unique biological projects. The publication record reflects sustained impact, with the platform and its major methodological extensions generating significant citations. Together, the usage and citation records show that Auto-Rickshaw has functioned both as a software service and as a crystallographic methodology that influenced automated experimental phasing, MR-assisted phasing and beamline-oriented structure solution.
8.1. Broader scientific and biopharmaceutical impact
The contribution of Auto-Rickshaw extended beyond methods development and individual proof-of-principle applications. During the structural genomics era, automated phasing was important for converting the large numbers of diffraction datasets generated by high-throughput programmes into interpretable electron-density maps and initial models. The MRSAD protocol, for example, was developed and evaluated using ten Joint Center for Structural Genomics cases, demonstrating how weak anomalous information could complement incomplete molecular-replacement models (Panjikar et al., 2009). Its use was not confined to these test cases. A search of the Protein Data Bank for depositions that explicitly record Auto-Rickshaw as structure-solution or phasing software identifies at least 282 entries at the time of writing. These include structures produced by the Joint Center for Structural Genomics, Midwest Center for Structural Genomics, Northeast Structural Genomics Consortium, Structural Genomics Consortium and Center for Structural Genomics of Infectious Diseases. Because software provenance has not always been recorded comprehensively in PDB depositions, this number is likely to represent a lower bound.
The deposited structures and their associated publications demonstrate a wide biological range. Applications in infectious-disease research include the Ebola virus interferon antagonist VP24, Marburg virus VP40, the Zika virus NS5 methyltransferase, a Legionella superbinding SH2 domain and the ferric preacinetobactin receptor BauA from the opportunistic pathogen Acinetobacter baumannii (Zhang et al., 2012; Oda et al., 2016; Coloma et al., 2016; Kaneko et al., 2018; Moynié et al., 2018). Other examples include proteins involved in bacterial signalling and biofilm formation, DNA and RNA recognition, restriction-endonuclease activity, chromosome organization and membrane transport (Schlundt et al., 2017; Tamulaitiene et al., 2014, 2017, 2019; Schneider et al., 2019; Chen et al., 2021). The corresponding PDB entries show that Auto-Rickshaw was applied to targets ranging from small soluble enzymes to multidomain, membrane-associated and protein–nucleic acid systems.
Several applications are also directly relevant to therapeutic discovery. Structures determined with the assistance of Auto-Rickshaw have supported the analysis of parasite-specific proteins and drug targets, including Plasmodium profilin, plasmepsins IX and X, Helicobacter pylori hypoxanthine–guanine–xanthine phosphoribosyltransferase and prolyl-tRNA synthetase complexes investigated as potential inhibitors of toxoplasmosis (Kursula et al., 2008; Hodder et al., 2022; Keough et al., 2021; Yogavel et al., 2023). The wider enzyme portfolio includes proteins involved in amino-acid and cofactor biosynthesis, plant metabolism and natural-product biosynthesis. These examples illustrate how automated structure solution can provide the structural starting point for mechanistic interpretation, target validation and structure-guided ligand development.
The scientific impact of Auto-Rickshaw was therefore not limited to the individual algorithms assembled within the pipeline. By providing experimental and hybrid phasing through a web-accessible workflow, checking alternative solution strategies automatically and returning interpretable maps and models without requiring users to master each underlying program, it lowered practical and computational barriers for crystallographers who were not specialists in phasing. This principle continues in work adapting automated structure-determination approaches to protein–ligand analysis and fragment-based drug-discovery pipelines (Khandokar et al., 2023).
9. Evolution: MRSAD, RIP, S-SAD and multi-dataset workflows
The history of Auto-Rickshaw after its initial release is a history of adaptation to crystallographic practice. As experimental methods evolved and user needs changed, the platform incorporated additional protocols and refinements that extended it beyond the initial SAD, MAD, SIRAS and MR workflows.
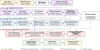
The development of MRSAD was one of the most significant extensions of Auto-Rickshaw (Panjikar et al., 2009). The combined MR/MRSAD workflow is illustrated in Fig. 3. The conceptual basis for combining molecular-replacement phases with weak anomalous information had been demonstrated earlier by Schuermann and Tanner, who showed that sulfur anomalous differences collected at Cu Kα wavelength could help overcome model bias in MR structure determination (Schuermann & Tanner, 2003). The Auto-Rickshaw implementation extended this idea into an automated decision-driven workflow. It responded to the increasing availability of homologous structures and to the practical reality that many molecular-replacement solutions are correct but only partially useful. A search model may provide enough phase information to guide density modification, but not enough to produce an interpretable map or reliable automated model building. Conversely, the anomalous signal may be too weak to solve the structure independently. By combining imperfect model-derived phases with anomalous differences, MRSAD formalized a hybrid strategy in which the two sources of phase information reinforce one another within an automated workflow.
|
Figure 3
MR/MRSAD phasing protocol in Auto-Rickshaw. The workflow combines a molecular-replacement branch, in which a search model is used to obtain and validate an MR solution, with an experimental anomalous, isomorphous or RIP branch, in which signal analysis and difference-map interpretation identify useful experimental sites. The MR-derived phases or partial model and the experimental sites are combined in the MR-assisted experimental phasing core, followed by dual-space fragment phasing, phase combination, density modification, model building and refinement. |
A demanding application of this strategy was the B–C protein complex from the Yersinia entomophaga ABC toxin system, originally reported as a large 2.5 Å resolution crystal structure in Nature (Busby et al., 2013). A subsequent methodological analysis described how cross-crystal averaging and Auto-Rickshaw MRSAD were combined to solve the 4354-amino-acid structure under challenging phasing conditions (Busby et al., 2016).
Radiation-induced phasing (RIP), molecular-replacement-assisted RIP (MRRIP) and UV-induced RIP represented another form of methodological expansion. The conceptual basis for RIP was established by Ravelli and coworkers, who showed that specific radiation damage could be used as a source of phase information for solving macromolecular crystal structures (Ravelli et al., 2003). This concept was subsequently extended to ultraviolet-induced structural changes, where UV exposure could generate site-specific differences suitable for phasing (Nanao & Ravelli, 2006). Later work showed that ultraviolet radiation could also generate useful, site-specific intensity changes in selenomethionine-containing protein crystals while retaining sufficient isomorphism for phasing. In particular, UV-induced cleavage around selenium sites enabled UV-RIP phasing of SeMet proteins (Panjikar et al., 2011), and related UV-RIPAS experiments combined UV-induced isomorphous differences with anomalous signal to produce phase sets comparable to those from conventional MAD approaches (de Sanctis et al., 2011). These protocols exploited radiation damage not only as a limitation of data collection but as a possible phasing signal. This inversion of perspective, turning an experimental complication into useful crystallographic information, fits naturally with the Auto-Rickshaw philosophy of exploring multiple possible routes to structure solution.
A subsequent structure determination of snakin-1 provides a useful example of RIP phasing in practice (Yeung et al., 2016). In this case, molecular replacement, ab initio attempts and an Auto-Rickshaw MAD strategy were unsuccessful, but radiation-damage differences between datasets enabled the automated RIP protocol of Auto-Rickshaw to solve the structure. Auto-Rickshaw then produced a largely traced model after SHELXE phasing, ARP/wARP model building and further density modification, as described in the supporting information to Yeung et al. (2016).
Sulfur-SAD also became increasingly important as synchrotron beamlines, detectors and data-collection strategies improved. Native sulfur phasing is attractive because it avoids derivatization or selenomethionine labelling, but it requires accurate measurement of weak anomalous differences. An important early demonstration was provided by Dauter and coworkers, who showed that the anomalous signal from S atoms could be used as a practical tool for protein structure solution, including lysozyme data collected at Cu Kα wavelength near 1.54 Å (Dauter et al., 1999). Automation can help by consistently evaluating the signal strength, testing appropriate resolution limits and applying robust downstream phasing procedures.
Multi-dataset workflows were another important area of development. Fast pixel-array detectors, brighter microfocus beamlines and automated sample changers made it practical to collect many partial or complete datasets from one or more crystals. Useful signal may be distributed across multiple crystals, multiple sweeps or multiple related datasets, so automated systems must manage not only a single set of reflections, but also the selection, comparison and combination of datasets. This motivated multiple-dataset modes in Auto-Rickshaw, including approaches in which datasets were selected, clustered or combined according to unit-cell similarity, isomorphism, radiation damage and anomalous signal before being passed into SAD, SIRAS or MAD phasing workflows (Panjikar et al., 2015, 2017). These developments extended the original single-job structure-solution concept towards beamline-scale data evaluation and automated selection of useful datasets.
10. Current status and outlook of the Auto-Rickshaw web service
The original EMBL Hamburg Auto-Rickshaw web service remained available for many years and supported a large number of external users, jobs and structure-solution projects. After the relocation of the principal developer from EMBL Hamburg to the Australian Synchrotron in June 2011, the public web service continued to be maintained remotely while further beamline-focused development proceeded at the Australian Synchrotron.
In August 2023, the public EMBL Hamburg web service was discontinued. Sustaining such a structural biology-oriented computational web service required a specialized combination of software-development, systems-administration, crystallographic and beamline expertise. By 2023, this complete support structure was no longer available locally at EMBL Hamburg. The discontinuation should therefore be understood as a change in service hosting and support capacity, rather than as a loss of scientific value or technical relevance of the pipeline.
At present, Auto-Rickshaw is available for macromolecular crystallography users at the Australian Synchrotron MX beamlines. By 2021, it had been installed on the ASCI cluster using Docker and Kubernetes for high-throughput job launching, enabling scalable execution from MX beamline computers through command-line operation, a web-based graphical interface and automatic data-processing workflows (Panjikar, 2021). This deployment also extended the use of the platform towards multiple-dataset evaluation, ligand-binding analysis and fragment-screening workflows.
A future goal is to identify a sustainable hosting model through which Auto-Rickshaw can be made available in its fullest capacity to a wider structural biology community. Such a model would need to balance open scientific access with modern requirements for cybersecurity, software maintenance, user management, licencing of external crystallographic programs and reproducible deployment. The recent ASWEBRICK and Docker-based developments are practical steps towards that goal.
11. From Auto-Rickshaw to ASWEBRICK: secure and reproducible deployment
A recent stage in the evolution of Auto-Rickshaw is the transition from a classical web-based structure-solution service to secured, containerized and beamline-integrated deployment. Automated structure-solution software increasingly forms part of an integrated beamline, data-management and decision-support environment rather than only a post-experiment analysis tool.
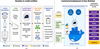
The evolution of Auto-Rickshaw deployment modes is summarized in Table 5 and illustrated in Fig. 4. The ASWEBRICK development represents a further step in this direction. Rather than treating Auto-Rickshaw only as a single public web service, ASWEBRICK emphasizes secure server operation, controlled user access and compatibility with institutional computing and cybersecurity requirements. In parallel, Docker-based deployment provides a portable environment in which the Auto-Rickshaw pipeline and pyAR (the Python scripting layer of Auto-Rickshaw) can run reproducibly while established crystallographic programs are accessed through controlled external software paths. This separation between the pipeline environment, user data and crystallographic software improves maintainability and makes the workflow more suitable for local, cluster-based and teaching deployments.
|
|||||||||||||||||||||||
|
Figure 4
Auto-Rickshaw as a bridge between beamline experiment and structure solution. (a) Conceptual beamline-to-model workflow. Diffraction data collected at a synchrotron beamline are processed and assessed before entering the Auto-Rickshaw decision-making layer, where appropriate phasing, density-modification, model-building and refinement strategies are selected and controlled. The resulting maps, models and diagnostic reports feed back into experimental decisions, including whether to collect additional data, change wavelength, test another crystal or proceed to model completion. (b) Modern containerized implementation of Auto-Rickshaw. Beamline or user data directories are mounted into the Docker container, separating read-only input data from read–write output directories. The Auto-Rickshaw and pyAR layers operate within a controlled software environment, while established crystallographic programs are accessed externally through a read-only software tree. This arrangement improves reproducibility, maintainability and deployment at synchrotron facilities. |
The inclusion of AlphaFold-derived or related search models is a natural extension, allowing modern model-prediction methods to enter a decision framework originally developed for experimental phasing and molecular replacement.
12. Auto-Rickshaw in the current landscape
The current structural biology landscape differs substantially from that of 2005. Detectors are faster, beamlines are more automated, remote and mail-in access are routine, fragment-screening and serial approaches generate large numbers of datasets, and computational infrastructure increasingly includes clusters, containers and cloud services. Molecular replacement has also become more powerful, aided by the growth of structural databases and, more recently, high-quality predicted models such as those generated by AlphaFold (Jumper et al., 2021).
Predicted structural models can now serve as search models within the molecular-replacement branch of the pipeline. When a suitable model is available, it can provide initial phases that may be combined with experimental information or passed through density-modification, model-building and refinement steps. Because this implementation has not yet been formally published, it is best regarded here as an emerging extension rather than as a fully documented protocol.
Recent developments in SHELXE reinforce this connection. Modern SHELXE workflows explicitly consider phases derived from partial or predicted models, and include approaches to reduce model bias during tracing and density modification (Usón & Sheldrick, 2024). This provides a conceptual link between AlphaFold-assisted molecular replacement, model-bias control and the earlier Auto-Rickshaw strategy of combining imperfect model-derived phases with experimental phasing information.
A further modern application area is fragment and ligand screening, where large numbers of protein–ligand datasets must be analysed rapidly. Work at the Australian Synchrotron has explored how automatic structure-determination approaches derived from Auto-Rickshaw can be adapted to streamline fragment-based drug-discovery pipelines and protein–ligand complex analysis (Khandokar et al., 2023).
These changes do not remove the need for experimental phasing pipelines. Difficult cases still involve weak anomalous signals, multiple datasets, radiation damage, partial models, low-resolution data and ambiguous maps. Modern pipelines may use containers, cloud execution, predicted models and machine-learning classifiers, but they still need to convert imperfect information into interpretable structural models.
The rise of AI-assisted crystallography gives this question renewed importance. Machine learning is already influencing several stages of the workflow, including crystal centring, diffraction-image classification and data reduction, model prediction, electron-density map interpretation and decision support (Ito et al., 2019; Ke et al., 2018; Jumper et al., 2021; Dialpuri et al., 2024; Vollmar & Evans, 2021). In this context, the evolution of Auto-Rickshaw illustrates how explicit domain knowledge, established algorithms and transparent diagnostics can be organized into a reliable decision-making workflow.
Looking ahead, an Auto-Rickshaw-style platform could combine rapid multi-dataset assessment with predicted search models, experimental anomalous evidence and automated model-bias checks, selecting among these sources according to their measured reliability rather than treating any one source as definitive (Jumper et al., 2021; Usón & Sheldrick, 2024). Machine-learning components could assist with classification and prioritization, while explicit crystallographic criteria and auditable intermediate results preserve scientific transparency. Secure containerized deployment could make the same validated workflow available at beamlines, institutional clusters and teaching facilities, and could support high-throughput applications such as fragment and ligand screening (Panjikar, 2021; Khandokar et al., 2023). The central future challenge is therefore not simply to add more algorithms, but to integrate new sources of structural information without losing reproducibility, diagnostics or expert oversight.
13. Lessons from Auto-Rickshaw's design philosophy
Several lessons emerge from the history of Auto-Rickshaw. Useful automation must be crystallographically informed: real diffraction data require conditional logic rather than a fixed list of commands. Robust defaults also matter, because many users need a scientifically reasonable first answer without losing expert control.
The platform also shows that software interoperability is a scientific capability. Auto-Rickshaw connected specialized programs through file conversion, input generation, output parsing, error handling, job control and result presentation. Finally, automation is most valuable when it remains transparent. Users need to know which route was taken, which programs were used, what intermediate statistics were obtained and why the calculation succeeded or failed.
14. Conclusions
Auto-Rickshaw was developed to guide experimental data through the decision landscape of macromolecular structure solution. Its historical importance lies not only in the number of phasing protocols it supported, but in the way it joined beamline needs, crystallographic algorithms, software engineering and expert decision making.
The integration of SHELXC, SHELXD and SHELXE gave Auto-Rickshaw a powerful experimental phasing core, while the broader pipeline connected this core to molecular replacement, density modification, automated model building and refinement. Extensions such as MRSAD, RIP, MRRIP, S-SAD and multiple-dataset workflows show how the platform evolved with the needs of the crystallographic community.
A short history of Auto-Rickshaw is therefore also a history of a larger transition: from manual execution of specialized programs towards adaptive, reproducible and beamline-aware structure-solution workflows. As structural biology moves towards unattended data collection, cloud computation, predicted models, electron-microscopy map interpretation and AI-assisted decision making, the principles embodied by Auto-Rickshaw remain relevant.
Footnotes
‡Retired group leader.
Acknowledgements
This review honours the scientific contributions of Professor George M. Sheldrick and the wider community of crystallographic software developers whose programs enabled automated structure-solution workflows. The historical development of Auto-Rickshaw was supported in part by the EC-funded BIOXHIT project (contract No. LHSG-CT-2003-503420) and the SPINE project (contract No. QLG2-CT-2002-00988), and benefited from the collaborative European structural biology environment at EMBL Hamburg. The authors gratefully acknowledge Venkataraman Parthasarathy for his important contributions to the IT infrastructure, web-server support and long-term operation of the Auto-Rickshaw service at EMBL Hamburg. The authors also acknowledge the many IT, systems-administration, software-support and beamline-computing colleagues at EMBL Hamburg and the Australian Synchrotron who made the long-term operation of Auto-Rickshaw possible. Their support in server maintenance, software deployment, user access, beamline integration and containerized computing has been essential to the evolution of Auto-Rickshaw from a public web service into modern synchrotron-linked and reproducible deployment environments. The authors acknowledge the assistance of ChatGPT, OpenAI in preparing schematic drafts from author-provided descriptions. All figures were subsequently checked, edited and ultimately approved by the authors. The authors also thank the many Auto-Rickshaw users, including synchrotron beamline visitors and remote web-service users, whose feedback and practical structure-solution problems helped guide the development and refinement of the platform. No specific funding was received for the preparation of this review.
Conflict of interest
The authors declare no conflicts of interest.
References
 Adams, P. D., Afonine, P. V., Bunkóczi, G., Chen, V. B., Davis, I. W., Echols, N., Headd, J. J., Hung, L.-W., Kapral, G. J., Grosse-Kunstleve, R. W., McCoy, A. J., Moriarty, N. W., Oeffner, R., Read, R. J., Richardson, D. C., Richardson, J. S., Terwilliger, T. C. & Zwart, P. H. (2010). Acta Cryst. D66, 213–221.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Australian Synchrotron (2011). AutoRickshaw at Your Service. https://archive.synchrotron.org.au/aussyncbeamlines/macromolecular-crystallography/highlights-mx/autorickshaw-at-your-service.
Google Scholar
BioSync (2026). Deposited PDB Structures for which Data were collected at European Synchrotron Facilities. https://biosync.rcsb.org/stats.do?stats_sec=RGNL&stats_focus_lvl=RGNL&stats_region=European.
Google Scholar
Brunzelle, J. S., Shafaee, P., Yang, X., Weigand, S., Ren, Z. & Anderson, W. F. (2003). Acta Cryst. D59, 1138–1144.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Busby, J. N., Lott, J. S. & Panjikar, S. (2016). Acta Cryst. D72, 182–191.
Web of Science
CrossRef
IUCr Journals
Google Scholar
Busby, J. N., Panjikar, S., Landsberg, M. J., Hurst, M. R. H. & Lott, J. S. (2013). Nature, 501, 547–550.
Web of Science
CrossRef
CAS
PubMed
Google Scholar
Chen, I., Pant, S., Wu, Q., Cater, R. J., Sobti, M., Vandenberg, R. J., Stewart, A. G., Tajkhorshid, E., Font, J. & Ryan, R. M. (2021). Nature, 591, 327–331.
CrossRef
PubMed
Google Scholar
Coloma, J., Jain, R., Rajashankar, K. R., García-Sastre, A. & Aggarwal, A. K. (2016). Cell. Rep. 16, 3097–3102.
Web of Science
CrossRef
CAS
PubMed
Google Scholar
Cowtan, K. (2006). Acta Cryst. D62, 1002–1011.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Dauter, Z., Dauter, M., de La Fortelle, E., Bricogne, G. & Sheldrick, G. M. (1999). J. Mol. Biol. 289, 83–92.
Web of Science
CrossRef
PubMed
CAS
Google Scholar
de La Fortelle, E. & Bricogne, G. (1997). Methods Enzymol. 276, 472–494.
CrossRef
CAS
PubMed
Web of Science
Google Scholar
de Sanctis, D., Tucker, P. A. & Panjikar, S. (2011). J. Synchrotron Rad. 18, 374–380.
Web of Science
CrossRef
IUCr Journals
Google Scholar
Dialpuri, J. S., Agirre, J., Cowtan, K. D. & Bond, P. S. (2024). Nucleic Acids Res. 52, e84.
CrossRef
PubMed
Google Scholar
Emsley, P., Lohkamp, B., Scott, W. G. & Cowtan, K. (2010). Acta Cryst. D66, 486–501.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Fernández, I. S., Ruíz-Dueñas, F. J., Santillana, E., Ferreira, P., Martínez, M. J., Martínez, Á. T. & Romero, A. (2009). Acta Cryst. D65, 1196–1205.
Web of Science
CrossRef
IUCr Journals
Google Scholar
Guss, J. M., Merritt, E. A., Phizackerley, R. P., Hedman, B., Murata, M., Hodgson, K. O. & Freeman, H. C. (1988). Science, 241, 806–811.
CrossRef
CAS
PubMed
Web of Science
Google Scholar
Hermes, C. (2004). AIP Conf. Proc. 705, 384–387.
CrossRef
Google Scholar
Hodder, A. N., Christensen, J., Scally, S., Triglia, T., Ngo, A., Birkinshaw, R. W., Bailey, B., Favuzza, P., Dietrich, M. H., Tham, W.-H., Czabotar, P. E., Lowes, K., Guo, Z., Murgolo, N., Lera Ruiz, M., McCauley, J. A., Sleebs, B. E., Olsen, D. B. & Cowman, A. F. (2022). Structure, 30, 947–961.
CrossRef
PubMed
Google Scholar
Holton, J. & Alber, T. (2004). Proc. Natl Acad. Sci. USA, 101, 1537–1542.
Web of Science
CrossRef
PubMed
CAS
Google Scholar
Ito, S., Ueno, G. & Yamamoto, M. (2019). J. Synchrotron Rad. 26, 1361–1366.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Jumper, J., Evans, R., Pritzel, A., Green, T., Figurnov, M., Ronneberger, O., Tunyasuvunakool, K., Bates, R., Žídek, A., Potapenko, A., Bridgland, A., Meyer, C., Kohl, S. A. A., Ballard, A. J., Cowie, A., Romera-Paredes, B., Nikolov, S., Jain, R., Adler, J., Back, T., Petersen, S., Reiman, D., Clancy, E., Zielinski, M., Steinegger, M., Pacholska, M., Berghammer, T., Bodenstein, S., Silver, D., Vinyals, O., Senior, A. W., Kavukcuoglu, K., Kohli, P. & Hassabis, D. (2021). Nature, 596, 583–589.
Web of Science
CrossRef
CAS
PubMed
Google Scholar
Kaneko, T., Stogios, P. J., Ruan, X., Voss, C., Evdokimova, E., Skarina, T., Chung, A., Liu, X., Li, L., Savchenko, A., Ensminger, A. W. & Li, S. S.-C. (2018). Nat. Commun. 9, 4549.
CrossRef
PubMed
Google Scholar
Ke, T.-W., Brewster, A. S., Yu, S. X., Ushizima, D., Yang, C. & Sauter, N. K. (2018). J. Synchrotron Rad. 25, 655–670.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Keough, D. T., Wun, S. J., Baszczyňski, O., Eng, W. S., Špaček, P., Panjikar, S., Naesens, L., Pohl, R., Rejman, D., Hocková, D., Ferrero, R. L. & Guddat, L. W. (2021). J. Med. Chem. 64, 5710–5729.
CrossRef
PubMed
Google Scholar
Khandokar, Y., Newton, S., Marshall, B., Caputo, A. T., Williamson, R., Panjikar, S. & Riboldi-Tunnicliffe, A. (2023). Acta Cryst. A79, C1054.
CrossRef
IUCr Journals
Google Scholar
Kroemer, M., Dreyer, M. K. & Wendt, K. U. (2004). Acta Cryst. D60, 1679–1682.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Kursula, I., Kursula, P., Ganter, M., Panjikar, S., Matuschewski, K. & Schüler, H. (2008). Structure, 16, 1638–1648.
Web of Science
CrossRef
PubMed
CAS
Google Scholar
Lamzin, V. S. & Wilson, K. S. (1993). Acta Cryst. D49, 129–147.
CrossRef
CAS
Web of Science
IUCr Journals
Google Scholar
Langer, G., Cohen, S. X., Lamzin, V. S. & Perrakis, A. (2008). Nat. Protoc. 3, 1171–1179.
Web of Science
CrossRef
PubMed
CAS
Google Scholar
Long, F., Vagin, A. A., Young, P. & Murshudov, G. N. (2008). Acta Cryst. D64, 125–132.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
McCoy, A. J., Grosse-Kunstleve, R. W., Adams, P. D., Winn, M. D., Storoni, L. C. & Read, R. J. (2007). J. Appl. Cryst. 40, 658–674.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Minor, W., Cymborowski, M., Otwinowski, Z. & Chruszcz, M. (2006). Acta Cryst. D62, 859–866.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Moynié, L., Serra, I., Scorciapino, M. A., Oueis, E., Page, M. G. P., Ceccarelli, M. & Naismith, J. H. (2018). eLife, 7, e42270.
PubMed
Google Scholar
Murshudov, G. N., Vagin, A. A. & Dodson, E. J. (1997). Acta Cryst. D53, 240–255.
CrossRef
CAS
Web of Science
IUCr Journals
Google Scholar
Nanao, M. H. & Ravelli, R. B. G. (2006). Structure, 14, 791–800.
Web of Science
CrossRef
PubMed
CAS
Google Scholar
Ness, S. R., de Graaff, R. A. G., Abrahams, J. P. & Pannu, N. S. (2004). Structure, 12, 1753–1761.
Web of Science
CrossRef
PubMed
CAS
Google Scholar
Oda, S.-I., Noda, T., Wijesinghe, K. J., Halfmann, P., Bornholdt, Z. A., Abelson, D. M., Armbrust, T., Stahelin, R. V., Kawaoka, Y. & Saphire, E. O. (2016). J. Virol. 90, 1839–1848.
CrossRef
PubMed
Google Scholar
Panjikar, S. (2010). Auto-Rickshaw. Presentation at the Macromolecular Crystallography School 2010, Madrid. https://www.xtal.iqfr.csic.es/MCS2010/Auto-Rickshaw-SP.pdf.
Google Scholar
Panjikar, S. (2021). Acta Cryst. A77, C529.
CrossRef
IUCr Journals
Google Scholar
Panjikar, S., Mayerhofer, H., Tucker, P. A., Mueller-Dieckmann, J. & de Sanctis, D. (2011). Acta Cryst. D67, 32–44.
Web of Science
CrossRef
IUCr Journals
Google Scholar
Panjikar, S., Parthasarathy, V., Lamzin, V. S., Weiss, M. S. & Tucker, P. A. (2005). Acta Cryst. D61, 449–457.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Panjikar, S., Parthasarathy, V., Lamzin, V. S., Weiss, M. S. & Tucker, P. A. (2009). Acta Cryst. D65, 1089–1097.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Panjikar, S., Weiss, M. & Lamzin, V. (2015). Acta Cryst. A71, s18.
CrossRef
IUCr Journals
Google Scholar
Panjikar, S., Weiss, M. & Lamzin, V. (2017). Acta Cryst. A73, C787.
Web of Science
CrossRef
IUCr Journals
Google Scholar
Pannu, N. S. & Read, R. J. (2004). Acta Cryst. D60, 22–27.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Pohl, E., González, A., Hermes, C. & van Silfhout, R. G. (2001). J. Synchrotron Rad. 8, 1113–1120.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Pohl, E., Ristau, U., Gehrmann, T., Jahn, D., Robrahn, B., Malthan, D., Dobler, H. & Hermes, C. (2004). J. Synchrotron Rad. 11, 372–377.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Ravelli, R. B. G., Leiros, H. S., Pan, B., Caffrey, M. & McSweeney, S. (2003). Structure, 11, 217–224.
Web of Science
CrossRef
PubMed
CAS
Google Scholar
Schlundt, A., Buchner, S., Janowski, R., Heydenreich, T., Heermann, R., Lassak, J., Geerlof, A., Stehle, R., Niessing, D., Jung, K. & Sattler, M. (2017). Sci. Rep. 7, 1051.
CrossRef
PubMed
Google Scholar
Schneider, T., Hung, L.-H., Aziz, M., Wilmen, A., Thaum, S., Wagner, J., Janowski, R., Müller, S., Schreiner, S., Friedhoff, P., Hüttelmaier, S., Niessing, D., Sattler, M., Schlundt, A. & Bindereif, A. (2019). Nat. Commun. 10, 2266.
CrossRef
PubMed
Google Scholar
Schneider, T. R. & Sheldrick, G. M. (2002). Acta Cryst. D58, 1772–1779.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Schuermann, J. P. & Tanner, J. J. (2003). Acta Cryst. D59, 1731–1736.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Sheldrick, G. M. (2008). Acta Cryst. A64, 112–122.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Sheldrick, G. M. (2010). Acta Cryst. D66, 479–485.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Tamulaitiene, G., Jovaisaite, V., Tamulaitis, G., Songailiene, I., Manakova, E., Zaremba, M., Grazulis, S., Xu, S. & Siksnys, V. (2017). Nucleic Acids Res. 45, 3547–3558.
PubMed
Google Scholar
Tamulaitiene, G., Manakova, E., Jovaisaite, V., Tamulaitis, G., Grazulis, S., Bochtler, M. & Siksnys, V. (2019). Nucleic Acids Res. 47, 997–1010.
CrossRef
PubMed
Google Scholar
Tamulaitiene, G., Silanskas, A., Grazulis, S., Zaremba, M. & Siksnys, V. (2014). Nucleic Acids Res. 42, 14022–14030.
CrossRef
PubMed
Google Scholar
Terwilliger, T. C. (2000). Acta Cryst. D56, 965–972.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Terwilliger, T. (2004). J. Synchrotron Rad. 11, 49–52.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Usón, I. & Sheldrick, G. M. (2018). Acta Cryst. D74, 106–116.
Web of Science
CrossRef
IUCr Journals
Google Scholar
Usón, I. & Sheldrick, G. M. (2024). Acta Cryst. D80, 4–15.
CrossRef
IUCr Journals
Google Scholar
Vagin, A. & Teplyakov, A. (1997). J. Appl. Cryst. 30, 1022–1025.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Vagin, A. & Teplyakov, A. (2010). Acta Cryst. D66, 22–25.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Vollmar, M. & Evans, G. (2021). Crystallogr. Rev. 27, 54–101.
Web of Science
CrossRef
Google Scholar
Vonrhein, C., Blanc, E., Roversi, P. & Bricogne, G. (2007). Methods Mol. Biol. 364, 215–230.
PubMed
CAS
Google Scholar
Winn, M. D., Ballard, C. C., Cowtan, K. D., Dodson, E. J., Emsley, P., Evans, P. R., Keegan, R. M., Krissinel, E. B., Leslie, A. G. W., McCoy, A., McNicholas, S. J., Murshudov, G. N., Pannu, N. S., Potterton, E. A., Powell, H. R., Read, R. J., Vagin, A. & Wilson, K. S. (2011). Acta Cryst. D67, 235–242.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Yeung, H., Squire, C. J., Yosaatmadja, Y., Panjikar, S., López, G., Molina, A., Baker, E. N., Harris, P. W. R. & Brimble, M. A. (2016). Angew. Chem. Int. Ed. 55, 7930–7933.
CrossRef
Google Scholar
Yogavel, M., Bougdour, A., Mishra, S., Malhotra, N., Chhibber-Goel, J., Bellini, V., Harlos, K., Laleu, B., Hakimi, M.-A. & Sharma, A. (2023). PLoS Pathog. 19, e1011124.
CrossRef
PubMed
Google Scholar
Zhang, A. P. P., Bornholdt, Z. A., Liu, T., Abelson, D. M., Lee, D. E., Li, S., Woods, V. L. J. & Saphire, E. O. (2012). PLoS Pathog. 8, e1002550.
CrossRef
PubMed
Google Scholar
This is an open-access article distributed under the terms of the Creative Commons Attribution (CC-BY) Licence, which permits unrestricted use, distribution, and reproduction in any medium, provided the original authors and source are cited.
| STRUCTURAL BIOLOGY |
access