Acta Crystallographica
Section E CRYSTALLOGRAPHIC COMMUNICATIONS


 menu


research communications
| CRYSTALLOGRAPHIC COMMUNICATIONS |
 access
of the pharmaceutical impurity 2-(4-chlorobenzoyl)pyridine

aDepartment of Chemistry, Vassar College, Poughkeepsie, NY 12604, USA
*Correspondence e-mail: [email protected]
The title compound, C12H8ClNO, is a pharmaceutical impurity and possible intermediate in the synthesis of carbinoxamine. The aromatic rings of the molecule twist relative to one another resulting in an interplanar angle of 38.1 (1)°. In the extended structure, the molecules are packed into rippled sheets with offset face-to-face stacking of each aromatic ring with the equivalent ring in the neighboring molecule.
CCDC reference: 2584887
1. Chemical context
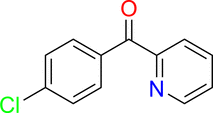
2-(4-Chlorobenzoyl)pyridine, C12H8ClNO (I), also known as carbinoxamine related compound A, is a known manufacturing impurity of the pharmaceutical ingredient carbinoxamine, 2-[(4-chlorophenyl)-pyridin-2-yl-methoxy]-N,N-dimethyl-ethanamine (C16H19ClN2O). The compound may be used to synthesize enantiopure alcohols, such as (S)-(4-chlorophenyl)(pyridin-2-yl)methanol, as precursors for the synthesis of carbinoxamine in the fungus Geotrichum candidum using a mutated acetophenone reductase enzyme that catalyzes the reduction of to (Tang et al., 2024 ). This enzyme belongs to the alcohol dehydrogenase (ADH) class, which catalyzes the reversible interconversion between ketones and alcohols. Carbinoxamine maleate, the maleic acid salt of carbinoxamine, is a first-generation H1-antihistamine used to treat allergy symptoms and may be synthesized via a Grignard reaction: para-chlorophenylmagnesium bromide, formed by the addition of para-bromochlorobenzene to magnesium in anhydrous ether, is treated with 2-pyridinealdehyde and subsequently with sodium and 2-dimethylaminoethyl chloride to produce an approximate 65% yield of the medication (Mansukhbhai & Das, 2021). Carbinoxamine maleate performs by binding to inactive H1 receptors belonging to G-protein-coupled receptors (GPCRs) found in cells and stabilizing them to remain inactive. This mechanism prevents the binding of the chemical signal histamine to the receptors and inhibits signal transduction pathways leading to allergic reactions (Leurs et al., 2002). As part of our studies in this area, we now describe the crystal structure of (I).
2. Structural commentary
Compound (I) crystallizes in space group P21/c with one molecule in the The molecular structure of (I) (Fig. 1) reveals a 4-chlorobenzoyl group, consisting of a benzene ring bearing a chlorine-atom substituent at the para position relative to the carbonyl group, which is attached to the carbon atom meta to the nitrogen atom of the pyridine ring. Individual molecules of (I) adopt a twisted conformation that minimizes intramolecular repulsion between the benzene and pyridine rings with an interplanar angle of 38.1 (1)° between the C1–C5/N1 and C7–C12 rings. The C5—C6 [1.506 (5) Å] and C6—C7 [1.506 (5) Å] bond lengths are almost identical and the N1—C5—C6—O1 torsion angle is −152.6 (4)°.
 | Figure 1 The molecular structure of (I) with displacement ellipsoids shown at the 50% probability level. |
3. Supramolecular features
In the extended structure, molecules of (I) assemble into rippled (100) sheets via weak C—H⋯O and C—H⋯Cl interactions (Fig. 2, Table 1). As expected, the C11—H11A⋯O1 interaction with a donor–acceptor distance of 3.406 (5) Å is shorter than the C2—H2A⋯Cl1 interaction with a corresponding distance of 3.633 (4) Å. The shortest Cl⋯Cl contact of 3.894 (6) Å is much longer than the sum of the van der Waals radii for chlorine (3.50 Å; Bondi, 1964). The sheets stack in such a manner that a slipped face-to-face π-stacking geometrical arrangement of the rings is present between equivalent rings within the molecules (Fig. 3). The π-stacking of the chlorobenzoyl rings is characterized by a centroid–centroid distance of 3.8943 (6) Å (the a unit-cell dimension) and a plane-to-centroid separation of 3.402 (3) Å, resulting in a ring shift of 1.895 (6) Å. The very similar π-stacking of the pyridine rings is characterized by the same centroid–centroid separation and a plane-to-centroid separation of 3.419 (3) Å, resulting in a ring shift of 1.865 (6) Å. In the overall packing (Fig. 4) there is also a longer C—H⋯N interaction (not shown) that interconnects the π-stacked sheets formed by the C—H⋯O and C—H⋯Cl interactions, with a C9—H9A⋯N1 donor–acceptor distance of 3.686 (5) Å.
| ||||||||||||||||||||||
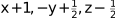 ; (ii)
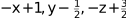 .
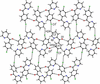 | Figure 2 A view of the weak hydrogen-bonded sheets in (I). Displacement ellipsoids are shown at the 50% probability level. Symmetry codes: (i) x + 1, −y +  , z - 1/2; (ii) −x + 1, y − , −z +  . |
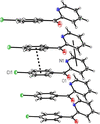 | Figure 3 A view of the face-to-face π-stacking geometrical arrangement in (I) with a dashed line indicating the chlorobenzoyl interaction and a solid line indicating the pyridine interaction. |
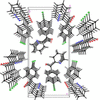 | Figure 4 The packing in (I) viewed down [100]. |
4. Database survey
One similar compound to (I) was found in the CSD version 6.00, April 2025.; Groom et al., 2016) with the CSD refcode CEYWOS (Syed et al., 1984), which consists of a 4-chlorobenzoyl group attached to a pyridine ring. However, in contrast to (I), the 4-chlorobenzoyl group in CEYWOS is attached to the carbon atom para to the nitrogen atom of the pyridine ring instead of the meta carbon atom. Both CEYWOS (space group P21) and (I) display similar weak intermolecular hydrogen bonding as well as π–π stacking interactions between the benzene and pyridine rings in parallel molecules. However, CEYWOS exhibits a greater twist angle of the benzoyl and pyridine rings relative to one another, resulting in an interplanar angle of 52.41° and a shortest Cl⋯Cl contact distance of 3.630 Å that is much closer to the sum of the van der Waals radii for chlorine.
5. Synthesis and crystallization
2-(4-Chlorobenzoyl)pyridine (95%) was purchased from Enamine, USA, and recrystallized by slow evaporation over 48 h at room temperature using a 1:1 solvent mixture of ethyl acetate/hexane.
6. Refinement
Crystal data, data collection and structure details are summarized in Table 2. Hydrogen atoms were included in calculated positions and refined using a riding model with C—H = 0.95 Å and Uiso(H) = 1.2Ueq(C).
|
Supporting information
CCDC reference: 2584887
contains datablocks global, I. DOI: https://doi.org/10.1107/S2056989026009175/hb8252sup1.cif
Structure factors: contains datablock I. DOI: https://doi.org/10.1107/S2056989026009175/hb8252Isup2.hkl
Supporting information file. DOI: https://doi.org/10.1107/S2056989026009175/hb8252Isup3.cml
| C12H8ClNO | F(000) = 448 |
| Mr = 217.64 | Dx = 1.469 Mg m−3 |
| Monoclinic, P21/c | Mo Kα radiation, λ = 0.71073 Å |
| a = 3.8943 (6) Å | Cell parameters from 9976 reflections |
| b = 10.8402 (15) Å | θ = 2.6–30.5° |
| c = 23.307 (3) Å | µ = 0.36 mm−1 |
| β = 90.573 (2)° | T = 125 K |
| V = 983.9 (2) Å3 | Plate, colourless |
| Z = 4 | 0.30 × 0.17 × 0.04 mm |
| Bruker APEXII CCD diffractometer | 3018 independent reflections |
| Radiation source: sealed X-ray tube, Bruker APEXII CCD | 2627 reflections with I > 2σ(I) |
| Graphite monochromator | Rint = 0.047 |
| Detector resolution: 8.3333 pixels mm-1 | θmax = 30.6°, θmin = 2.1° |
| φ and ω scans | h = −5→5 |
| Absorption correction: multi-scan (SADABS; Krause et al., 2015) | k = −15→15 |
| Tmin = 0.87, Tmax = 0.99 | l = −33→33 |
| 22981 measured reflections |
| Refinement on F2 | Primary atom site location: dual |
| Least-squares matrix: full | Secondary atom site location: difference Fourier map |
| R[F2 > 2σ(F2)] = 0.087 | Hydrogen site location: inferred from neighbouring sites |
| wR(F2) = 0.234 | H-atom parameters constrained |
| S = 1.19 | w = 1/[σ2(Fo2) + (0.0373P)2 + 7.3622P] where P = (Fo2 + 2Fc2)/3 |
| 3018 reflections | (Δ/σ)max < 0.001 |
| 136 parameters | Δρmax = 0.63 e Å−3 |
| 0 restraints | Δρmin = −0.91 e Å−3 |
Geometry. All esds (except the esd in the dihedral angle between two l.s. planes) are estimated using the full covariance matrix. The cell esds are taken into account individually in the estimation of esds in distances, angles and torsion angles; correlations between esds in cell parameters are only used when they are defined by crystal symmetry. An approximate (isotropic) treatment of cell esds is used for estimating esds involving l.s. planes. |
| x | y | z | Uiso*/Ueq | ||
| Cl1 | 0.1218 (3) | 0.36829 (10) | 0.89790 (4) | 0.0240 (3) | |
| O1 | 0.3127 (10) | 0.5625 (3) | 0.62865 (14) | 0.0306 (8) | |
| N1 | 0.3630 (9) | 0.2415 (3) | 0.61545 (14) | 0.0182 (6) | |
| C1 | 0.4401 (11) | 0.1573 (4) | 0.57571 (17) | 0.0209 (8) | |
| H1B | 0.375692 | 0.074018 | 0.582413 | 0.025* | |
| C2 | 0.6100 (11) | 0.1851 (4) | 0.52497 (18) | 0.0220 (8) | |
| H2A | 0.66341 | 0.121867 | 0.498278 | 0.026* | |
| C3 | 0.6995 (11) | 0.3065 (4) | 0.51413 (19) | 0.0247 (8) | |
| H3A | 0.810253 | 0.328332 | 0.47945 | 0.03* | |
| C4 | 0.6246 (11) | 0.3954 (4) | 0.55481 (17) | 0.0209 (8) | |
| H4A | 0.687546 | 0.479185 | 0.549092 | 0.025* | |
| C5 | 0.4537 (10) | 0.3588 (4) | 0.60456 (16) | 0.0167 (7) | |
| C6 | 0.3522 (11) | 0.4574 (4) | 0.64667 (17) | 0.0186 (7) | |
| C7 | 0.3001 (10) | 0.4297 (3) | 0.70928 (16) | 0.0159 (7) | |
| C8 | 0.1338 (10) | 0.5204 (3) | 0.74168 (17) | 0.0165 (7) | |
| H8A | 0.055346 | 0.593758 | 0.723501 | 0.02* | |
| C9 | 0.0826 (10) | 0.5041 (4) | 0.79999 (17) | 0.0193 (7) | |
| H9A | −0.027204 | 0.566012 | 0.82209 | 0.023* | |
| C10 | 0.1959 (10) | 0.3951 (4) | 0.82540 (16) | 0.0172 (7) | |
| C11 | 0.3722 (10) | 0.3054 (4) | 0.79449 (17) | 0.0174 (7) | |
| H11A | 0.458215 | 0.233698 | 0.813115 | 0.021* | |
| C12 | 0.4203 (10) | 0.3227 (4) | 0.73593 (17) | 0.0174 (7) | |
| H12A | 0.534994 | 0.261473 | 0.714114 | 0.021* |
| U11 | U22 | U33 | U12 | U13 | U23 | |
| Cl1 | 0.0307 (5) | 0.0248 (5) | 0.0165 (4) | −0.0023 (4) | 0.0050 (3) | 0.0009 (4) |
| O1 | 0.054 (2) | 0.0156 (14) | 0.0218 (15) | 0.0020 (14) | 0.0022 (14) | 0.0033 (12) |
| N1 | 0.0213 (16) | 0.0166 (15) | 0.0168 (15) | −0.0006 (12) | −0.0001 (12) | −0.0006 (12) |
| C1 | 0.0264 (19) | 0.0161 (17) | 0.0201 (18) | −0.0016 (15) | 0.0015 (15) | −0.0030 (14) |
| C2 | 0.0225 (19) | 0.025 (2) | 0.0184 (18) | 0.0022 (16) | 0.0024 (14) | −0.0043 (15) |
| C3 | 0.024 (2) | 0.029 (2) | 0.0208 (19) | 0.0014 (17) | 0.0040 (15) | 0.0023 (16) |
| C4 | 0.0223 (18) | 0.0207 (19) | 0.0198 (18) | −0.0013 (15) | 0.0011 (14) | 0.0029 (15) |
| C5 | 0.0184 (16) | 0.0159 (16) | 0.0156 (16) | −0.0017 (13) | −0.0031 (13) | −0.0002 (13) |
| C6 | 0.0233 (18) | 0.0147 (16) | 0.0176 (17) | −0.0021 (14) | −0.0022 (14) | 0.0005 (13) |
| C7 | 0.0181 (16) | 0.0132 (16) | 0.0163 (16) | −0.0027 (13) | −0.0016 (13) | 0.0002 (13) |
| C8 | 0.0163 (16) | 0.0123 (16) | 0.0210 (17) | −0.0001 (13) | −0.0007 (13) | −0.0003 (13) |
| C9 | 0.0200 (18) | 0.0177 (17) | 0.0204 (18) | 0.0012 (14) | 0.0038 (14) | −0.0018 (14) |
| C10 | 0.0190 (17) | 0.0178 (17) | 0.0149 (16) | −0.0024 (13) | 0.0004 (13) | −0.0001 (13) |
| C11 | 0.0198 (17) | 0.0142 (16) | 0.0181 (17) | 0.0015 (13) | −0.0030 (13) | 0.0008 (13) |
| C12 | 0.0189 (17) | 0.0144 (17) | 0.0188 (17) | 0.0014 (13) | −0.0004 (13) | −0.0022 (13) |
| Cl1—C10 | 1.741 (4) | C5—C6 | 1.506 (5) |
| O1—C6 | 1.224 (5) | C6—C7 | 1.506 (5) |
| N1—C1 | 1.337 (5) | C7—C12 | 1.394 (5) |
| N1—C5 | 1.345 (5) | C7—C8 | 1.403 (5) |
| C1—C2 | 1.394 (6) | C8—C9 | 1.387 (5) |
| C1—H1B | 0.95 | C8—H8A | 0.95 |
| C2—C3 | 1.385 (6) | C9—C10 | 1.392 (6) |
| C2—H2A | 0.95 | C9—H9A | 0.95 |
| C3—C4 | 1.386 (6) | C10—C11 | 1.395 (5) |
| C3—H3A | 0.95 | C11—C12 | 1.392 (5) |
| C4—C5 | 1.400 (5) | C11—H11A | 0.95 |
| C4—H4A | 0.95 | C12—H12A | 0.95 |
| C1—N1—C5 | 117.0 (3) | C12—C7—C8 | 119.9 (3) |
| N1—C1—C2 | 123.5 (4) | C12—C7—C6 | 123.3 (3) |
| N1—C1—H1B | 118.3 | C8—C7—C6 | 116.7 (3) |
| C2—C1—H1B | 118.3 | C9—C8—C7 | 120.7 (4) |
| C3—C2—C1 | 118.9 (4) | C9—C8—H8A | 119.7 |
| C3—C2—H2A | 120.6 | C7—C8—H8A | 119.7 |
| C1—C2—H2A | 120.6 | C8—C9—C10 | 118.5 (4) |
| C2—C3—C4 | 118.8 (4) | C8—C9—H9A | 120.8 |
| C2—C3—H3A | 120.6 | C10—C9—H9A | 120.8 |
| C4—C3—H3A | 120.6 | C9—C10—C11 | 121.8 (4) |
| C3—C4—C5 | 118.3 (4) | C9—C10—Cl1 | 120.0 (3) |
| C3—C4—H4A | 120.8 | C11—C10—Cl1 | 118.2 (3) |
| C5—C4—H4A | 120.8 | C12—C11—C10 | 119.0 (4) |
| N1—C5—C4 | 123.5 (4) | C12—C11—H11A | 120.5 |
| N1—C5—C6 | 118.5 (3) | C10—C11—H11A | 120.5 |
| C4—C5—C6 | 117.9 (4) | C11—C12—C7 | 120.0 (4) |
| O1—C6—C7 | 120.0 (4) | C11—C12—H12A | 120.0 |
| O1—C6—C5 | 118.0 (4) | C7—C12—H12A | 120.0 |
| C7—C6—C5 | 121.9 (3) | ||
| C5—N1—C1—C2 | 0.7 (6) | C5—C6—C7—C12 | 16.5 (6) |
| N1—C1—C2—C3 | −1.2 (7) | O1—C6—C7—C8 | 13.9 (6) |
| C1—C2—C3—C4 | 1.6 (7) | C5—C6—C7—C8 | −166.2 (4) |
| C2—C3—C4—C5 | −1.5 (6) | C12—C7—C8—C9 | −1.0 (6) |
| C1—N1—C5—C4 | −0.7 (6) | C6—C7—C8—C9 | −178.4 (4) |
| C1—N1—C5—C6 | 176.4 (4) | C7—C8—C9—C10 | −0.9 (6) |
| C3—C4—C5—N1 | 1.1 (6) | C8—C9—C10—C11 | 3.1 (6) |
| C3—C4—C5—C6 | −176.0 (4) | C8—C9—C10—Cl1 | −177.5 (3) |
| N1—C5—C6—O1 | −152.6 (4) | C9—C10—C11—C12 | −3.4 (6) |
| C4—C5—C6—O1 | 24.6 (6) | Cl1—C10—C11—C12 | 177.1 (3) |
| N1—C5—C6—C7 | 27.5 (6) | C10—C11—C12—C7 | 1.5 (6) |
| C4—C5—C6—C7 | −155.3 (4) | C8—C7—C12—C11 | 0.6 (6) |
| O1—C6—C7—C12 | −163.4 (4) | C6—C7—C12—C11 | 177.8 (4) |
| D—H···A | D—H | H···A | D···A | D—H···A |
| C2—H2A···Cl1i | 0.95 | 2.96 | 3.633 (4) | 129 |
| C11—H11A···O1ii | 0.95 | 2.46 | 3.406 (5) | 173 |
| Symmetry codes: (i) x+1, −y+1/2, z−1/2; (ii) −x+1, y−1/2, −z+3/2. |
Acknowledgements
This work was supported by Vassar College. X-ray facilities were provided by the US National Science Foundation (grant Nos. 0521237 and 0911324 to JT).
Funding information
Funding for this research was provided by: National Science Foundation, Directorate for Mathematical and Physical Sciences (grant No. 0521237 to Joseph M. Tanski; award No. 0911324 to Joseph M. Tanski).
References
 Bondi, A. (1964). J. Phys. Chem. 68, 441–451. CrossRef CAS Web of Science Google Scholar
Bruker (2013). SAINT and APEX3. Bruker AXS Inc., Madison, Wisconsin, USA. Google Scholar
Dolomanov, O. V., Bourhis, L. J., Gildea, R. J., Howard, J. A. K. & Puschmann, H. (2009). J. Appl. Cryst. 42, 339–341. Web of Science CrossRef CAS IUCr Journals Google Scholar
Groom, C. R., Bruno, I. J., Lightfoot, M. P. & Ward, S. C. (2016). Acta Cryst. B72, 171–179. Web of Science CrossRef IUCr Journals Google Scholar
Krause, L., Herbst-Irmer, R., Sheldrick, G. M. & Stalke, D. (2015). J. Appl. Cryst. 48, 3–10. Web of Science CSD CrossRef ICSD CAS IUCr Journals Google Scholar
Leurs, R., Church, M. K. & Taglialatela, M. (2002). Clin. Exp. Allergy 32, 489–498. CrossRef PubMed CAS Google Scholar
Macrae, C. F., Sovago, I., Cottrell, S. J., Galek, P. T. A., McCabe, P., Pidcock, E., Platings, M., Shields, G. P., Stevens, J. S., Towler, M. & Wood, P. A. (2020). J. Appl. Cryst. 53, 226–235. Web of Science CrossRef CAS IUCr Journals Google Scholar
Mansukhbhai, S. A. & Das, A. K. (2021). J. Pharm. Res. 20, 21–26. CrossRef Google Scholar
Sheldrick, G. M. (2008). Acta Cryst. A64, 112–122. Web of Science CrossRef CAS IUCr Journals Google Scholar
Sheldrick, G. M. (2015a). Acta Cryst. A71, 3–8. Web of Science CrossRef IUCr Journals Google Scholar
Sheldrick, G. M. (2015b). Acta Cryst. C71, 3–8. Web of Science CrossRef IUCr Journals Google Scholar
Syed, A., Foss, L. I. & Stevens, E. D. (1984). Acta Cryst. C40, 834–836. CrossRef CAS IUCr Journals Google Scholar
Tang, Z., Takagi, Y., Koesoema, A. A. & Matsuda, T. (2024). Appl. Microbiol. Biotechnol. 108, 545. CrossRef PubMed Google Scholar
This is an open-access article distributed under the terms of the Creative Commons Attribution (CC-BY) Licence, which permits unrestricted use, distribution, and reproduction in any medium, provided the original authors and source are cited.
| CRYSTALLOGRAPHIC COMMUNICATIONS |