Acta Crystallographica
Section E CRYSTALLOGRAPHIC COMMUNICATIONS


 menu


research communications
| CRYSTALLOGRAPHIC COMMUNICATIONS |
 access
Zn(AsO3)2, a unique structure among arsenates with composition MIIAsV2O6

aTU Wien, X-Ray Centre, Getreidemarkt 9/E057, 1060 Vienna, Austria, and bTU Wien, Institute for Chemical Technologies and Analytics, Division of Applied Solid State Chemistry, Getreidemarkt 9/E164-05-1, 1060 Vienna, Austria
*Correspondence e-mail: [email protected]
Zinc bis[catena-arsenate(V)] or catena-poly[zinc-di-μ-arsenato(V)-κ4O:O], [Zn(AsO3)2]n, is isotypic with the low-temperature modification of Zn(PO3)2. The asymmetric unit comprises one Zn atom situated on a twofold rotation axis and one As and three O atoms in general positions. In the crystal structure, the principal building units, ZnO6 octahedra and AsO4 tetrahedra, are condensed into 1∞[ZnO4/2O2/1] zigzag chains and (AsO3)2 polyarsenate chains with a periodicity of two AsO4 tetrahedra, respectively. The two types of chains extend parallel to [001] and are fused into a compact framework structure. Compared with divalent transition metal arsenates and other arsenates of the formula MIIAs2O6, Zn(AsO3)2 is unique in that arsenic is present exclusively in a tetrahedral coordination and not in a mixed tetrahedral/octahedral or purely octahedral coordination.
Keywords: crystal structure; structural comparison; isotypism; polyarsenate chain.
CCDC reference: 2585628
1. Chemical context
The zinc arsenate Zn(AsO3)2 has been reported as one of the phases existing in the pseudo-binary system ZnO–As2O5 (Kasenov et al., 1996 ). Whereas for the other phases reported in this system complete crystal structure determinations have been carried out in the meantime (details are given in the database survey), for Zn(AsO3)2 only unit-cell parameters were given on basis of a powder X-ray diffraction study: orthorhombic with a = 8.625, b = 9.035, c = 12.494 Å, V = 933 Å3 (Mustafin & Kasenov, 1994, 1996). Recent results of crystal structure prediction methods claimed the possible existence of Zn(AsO3)2 (or as stated there of ZnAs2O6). Whereas one study states that this phase has an existence probability of 0.9105 with a polar structure (Gake et al., 2025; data given in the supporting information) the other states that it crystallizes in the centrosymmetric PbSb2O6 structure type with trigonal symmetry (Griesemer et al., 2025; supplementary material of this article: OQMD ID 1348395).
We sought to investigate these conflicting claims and carried out experiments on the crystal growth of Zn(AsO3)2. Using a reaction similar to that described by Kasenov et al. (1996), we were able to isolate single crystals of Zn(AsO3)2 (I) and report here on the results of the crystal structure analysis.
2. Structural commentary
Our study does not confirm any of the previous statements made regarding the orthorhombic or possible crystal structures. Compound (I) crystallizes in the monoclinic system in the centrosymmetric space group C2/c and is isotypic with the corresponding phosphate, i.e. low-temperature Zn(PO3)2 (Averbuch-Pouchot et al., 1983). The unit cell of Zn(AsO3)2 reported by Mustafin & Kasenov (1994, 1996) has a doubled volume compared to the unit cell determined here, however, with no apparent metrical relation beyond this.
In the crystal structure of Zn(AsO3)2, the Zn atom is located on a twofold rotation axis (multiplicity 4, Wyckoff letter e), and the As and the three O atom sites on general positions (8f) of space group C2/c. The zinc atom in (I) exhibits a distorted octahedral coordination environment with an average Zn—O bond length of 2.090 Å (Table 1), in good agreement with the literature value of 2.110 (86) Å derived from 193 [ZnO6] coordination polyhedra (Gagné & Hawthorne, 2020). The [ZnO6] octahedron shares two of its edges with neighbouring octahedra, forming 1∞[ZnO4/2O2/1] zigzag-chains extending parallel to [001] (Fig. 1a). The arsenate tetrahedron condenses into a polyarsenate chain with a periodicity of two tetrahedra (Fig. 1b). The bond length distribution (Table 1) in this chain follows the usual pattern, with significantly longer As—O distances to the bridging oxygen atoms (O1, average 1.735 Å) than to the terminal O atoms (O2, O3; average 1.646 Å). The mean average As—O bond length within the AsO4 tetrahedron (1.690 Å) complies with the literature value [1.687 (27) Å from 508 arsenate tetrahedra; Gagné & Hawthorne, 2018]. The terminal oxygen atoms of the polyarsenate chain are also the ones that bind to the zinc atoms in adjacent zinc oxide chains, so that the polyarsenate chains are aligned in the same direction as the zinc oxide chains, which leads to a very compact packing in this type of structure (Durif, 1995). In the crystal structure of (I) (Fig. 2), each zinc oxide chain is surrounded by six polyarsenate chains, and each polyarsenate chain is surrounded by three zinc oxide chains, corresponding to the 1:2 ratio of Zn:As in the formula.
| ||||||||||||||||||||||
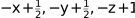 ; (ii)
 ; (iii)
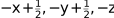 ; (iv)
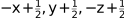 .
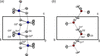 | Figure 1 (a) One 1∞[ZnO4/2O2/1] zigzag chain, and (b) one polyarsenate chain crossing the unit cell of (I), as shown in projections down [  00]. Displacement ellipsoids are drawn at the 97% probability level. [Symmetry codes: (i) x +  , y + , z; (ii) −x + 1, −y + 1, −z + 1; (iii) −x + 1, y, −z + ; (iv) x, −y + 1, z − ; (v) −x + , y + , −z + ; (vi) −x + , −y + , −z.] |
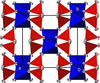 | Figure 2 The crystal structure of (I) in polyhedral representation (ZnO6 octahedra blue, AsO4 tetrahedra red), viewed down [00 ]. Displacement ellipsoids are drawn at the 97% probability level. |
The structure model was confirmed using bond-valence-sum calculations (Brown, 2002) performed with the ECoN21 program (Ilinca, 2022). The calculated bond-valence sums (Table 2) are close to the expected values of 2, 5 and 2 valence units for Zn, As and O atoms, respectively, and the global instability index (GII) of 0.11 valence units indicates a stable and not particularly strained crystal structure (Salinas-Sanchez et al., 1992; Brown, 2009).
| ||||||||||||||||||||||||||
Using the compstru program (de la Flor et al., 2016) that is available at the Bilbao Crystallographic Server (Aroyo et al., 2006), a quantitative comparison of the isotypic structures of low-temperature Zn(PO3)2 and Zn(AsO3)2 was made. The arithmetic mean of the distance between paired atoms is 0.1148 Å, and the absolute distances between paired atoms are 0.0925 Å for Zn1, 0.0598 Å for As1/P1, 0.0901 Å for O1, 0.1146 Å for O2, and the largest distance of 0.2057 Å for O3. The latter O atoms are those which, in both structures, have both the shortest Zn—O and the shortest X—O (X = As, P) bond lengths. The degree of (0.0264), and the low value for the measure of similarity (0.023) indicate a high degree of similarity for the two crystal structures.
3. Database survey
The Inorganic Database (ICSD, data release 2025.1; Zagorac et al., 2019) was used for the database search. Known structurally characterized phases in the ZnO–As2O5 system include the high- and low-temperature modifications of tetrazinc bis(arsenate(V)) oxide, Zn4(AsO4)2O (Frerichs et al., 1996a), dimorphic trizinc orthoarsenate, Zn3(AsO4)2 (Frerichs et al., 1996b), and the pyroarsenate Zn2As2O7 with three modifications existing at different temperatures, one of which is commensurately and one is incommensurately modulated (Weil & Stöger, 2010).
As already mentioned, Zn(AsO3)2 is isotypic with Zn(PO3)2, in its low-temperature modification (Averbuch-Pouchot et al., 1983). The high-temperature modification of Zn(PO3)2 can be obtained by quenching from the recrystallized melt and contains a similar polyphosphate chain but in contrast to the low-temperature modification with two unique zinc cations in tetrahedral coordination (Weil, 2004). According to the information on the system ZnO–As2O5 provided by Kasenov et al. (1996), there are no indications of a possible high-temperature form of Zn(AsO3)2. Other M(PO3)2 polyphosphates isotypic with low-temperature Zn(PO3)2 and Zn(AsO3)2 can only be prepared by using high pressure (∼80 kbar) and temperatures (∼1270 K) and are reported for M = Ni, Mg, Cu, Co, Fe, Mn, and Cd (Bagieu-Beucher et al., 1976).
Zn(AsO3)2 is the only arsenate phase with an M:AsV ratio of 1:2 where solely tetrahedral AsO4 units are present in the polymeric arsenate anion. Except for M = Ba with a catena-arsenate anion consisting of AsO4 tetrahedra and AsO6 octahedra (Weil, 2016), all other MAs2O6 phases crystallize in the PbSb2O6 type of structure (Hill, 1987) in the trigonal space group type P 1m. This structure type is also referred to as the rosiaite structure type after the mineral of this name (Basso et al., 1996) and is based on a hexagonal array of O atoms with the As atom octahedrally coordinated in form of honeycomb layers. This structure type is realized for M = Ni, Co, Mn (Nakua & Greedan, 1995), Cd (Weil, 2001), Hg (Weil, 2000), Pd (Orosel & Jansen, 2006), Ca, Pb (Losilla et al., 1995), and Sr. The strontium phase is dimorphic and as well as in a rosiaite-type structure also crystallizes in a hexagonal superstructure with a doubled unit cell (Weil et al., 2009).
4. Synthesis and crystallization
Crystal growth procedures were carried out in sealed and evacuated silica ampoules using commercially available chemicals as received. A molar 1:1 mixture of ZnO (≥ 99%, Carl Roth GmbH) and As2O5 [99.9% (metals basis), abcr] was heated at 750 K for about 336 h. X-ray powder diffraction of the bulk product revealed Zn(AsO3OH)·H2O (mineral name koritnigite; Keller et al., 1980) and Zn(AsO3)2 as the main phases in an approximate ratio of 1:2. From this mixture, yellowish crystals of Zn(AsO3)2 were isolated for the single-crystal X-ray diffraction study. The presence of the hydrous phase Zn(AsO3OH)·H2O indicates that the hygroscopic starting material As2O5 was indeed not free of water.
5. Refinement
Crystal data, data collection and structure details are summarized in Table 3. Atomic coordinates and labelling were adapted from the isotypic low-temperature phase of Zn(PO3)2 (Averbuch-Pouchot et al., 1983).
|
Supporting information
CCDC reference: 2585628
contains datablock I. DOI: https://doi.org/10.1107/S2056989026009308/hb8254sup1.cif
Structure factors: contains datablock I. DOI: https://doi.org/10.1107/S2056989026009308/hb8254Isup2.hkl
| [Zn(AsO3)2] | F(000) = 576 |
| Mr = 311.21 | Dx = 4.520 Mg m−3 |
| Monoclinic, C2/c | Mo Kα radiation, λ = 0.71073 Å |
| a = 10.1061 (10) Å | Cell parameters from 5088 reflections |
| b = 9.0308 (8) Å | θ = 3.1–32.8° |
| c = 5.2801 (5) Å | µ = 19.68 mm−1 |
| β = 108.374 (7)° | T = 300 K |
| V = 457.33 (8) Å3 | Shard, yellow |
| Z = 4 | 0.04 × 0.02 × 0.01 mm |
| Stoe Stadivari diffractometer | 797 independent reflections |
| Radiation source: Axo_Mo | 770 reflections with I > 2σ(I) |
| Graded multilayer mirror monochromator | Rint = 0.017 |
| Detector resolution: 13.33 pixels mm-1 | θmax = 32.8°, θmin = 3.1° |
| rotation method, ω scans | h = −14→15 |
| Absorption correction: multi-scan (LANA; Koziskova et al., 2016) | k = −12→13 |
| Tmin = 0.492, Tmax = 0.654 | l = −7→4 |
| 3846 measured reflections |
| Refinement on F2 | Primary atom site location: isomorphous structure methods |
| Least-squares matrix: full | w = 1/[σ2(Fo2) + (0.0127P)2 + 0.509P] where P = (Fo2 + 2Fc2)/3 |
| R[F2 > 2σ(F2)] = 0.010 | (Δ/σ)max = 0.001 |
| wR(F2) = 0.028 | Δρmax = 0.40 e Å−3 |
| S = 1.13 | Δρmin = −0.36 e Å−3 |
| 797 reflections | Extinction correction: SHELXL (Sheldrick, 2015), Fc*=kFc[1+0.001xFc2λ3/sin(2θ)]-1/4 |
| 43 parameters | Extinction coefficient: 0.0035 (2) |
| 0 restraints |
Geometry. All esds (except the esd in the dihedral angle between two l.s. planes) are estimated using the full covariance matrix. The cell esds are taken into account individually in the estimation of esds in distances, angles and torsion angles; correlations between esds in cell parameters are only used when they are defined by crystal symmetry. An approximate (isotropic) treatment of cell esds is used for estimating esds involving l.s. planes. |
| x | y | z | Uiso*/Ueq | ||
| Zn1 | 0.000000 | 0.10801 (2) | 0.250000 | 0.00841 (6) | |
| As1 | 0.21468 (2) | 0.40539 (2) | 0.24523 (3) | 0.00640 (5) | |
| O1 | 0.36157 (10) | −0.00460 (12) | 0.04492 (19) | 0.0121 (2) | |
| O2 | 0.38566 (10) | 0.42724 (11) | 0.35206 (19) | 0.00870 (17) | |
| O3 | 0.15428 (11) | 0.23634 (11) | 0.1990 (2) | 0.0132 (2) |
| U11 | U22 | U33 | U12 | U13 | U23 | |
| Zn1 | 0.00662 (11) | 0.00985 (11) | 0.00828 (10) | 0.000 | 0.00168 (8) | 0.000 |
| As1 | 0.00579 (7) | 0.00774 (8) | 0.00560 (7) | −0.00148 (4) | 0.00169 (5) | −0.00034 (4) |
| O1 | 0.0078 (4) | 0.0209 (5) | 0.0074 (4) | 0.0001 (4) | 0.0021 (3) | 0.0056 (4) |
| O2 | 0.0057 (4) | 0.0118 (4) | 0.0077 (4) | −0.0008 (3) | 0.0008 (3) | 0.0008 (3) |
| O3 | 0.0139 (5) | 0.0094 (4) | 0.0172 (5) | −0.0052 (4) | 0.0061 (4) | −0.0020 (4) |
| Zn1—O3 | 2.0276 (10) | Zn1—O2v | 2.1640 (10) |
| Zn1—O3i | 2.0276 (10) | As1—O3 | 1.6336 (10) |
| Zn1—O2ii | 2.0786 (10) | As1—O2 | 1.6518 (10) |
| Zn1—O2iii | 2.0786 (10) | As1—O1vi | 1.7319 (10) |
| Zn1—O2iv | 2.1640 (10) | As1—O1vii | 1.7379 (10) |
| O3—Zn1—O3i | 110.28 (6) | O2iii—Zn1—O2v | 88.05 (3) |
| O3—Zn1—O2ii | 99.15 (4) | O2iv—Zn1—O2v | 82.06 (5) |
| O3i—Zn1—O2ii | 90.92 (4) | O3—As1—O2 | 117.65 (5) |
| O3—Zn1—O2iii | 90.92 (4) | O3—As1—O1vi | 108.08 (5) |
| O3i—Zn1—O2iii | 99.15 (4) | O2—As1—O1vi | 111.48 (5) |
| O2ii—Zn1—O2iii | 162.38 (6) | O3—As1—O1vii | 108.20 (5) |
| O3—Zn1—O2iv | 163.24 (4) | O2—As1—O1vii | 110.65 (5) |
| O3i—Zn1—O2iv | 84.54 (4) | O1vi—As1—O1vii | 99.15 (3) |
| O2ii—Zn1—O2iv | 88.05 (3) | As1vi—O1—As1v | 130.01 (6) |
| O2iii—Zn1—O2iv | 78.64 (4) | As1—O2—Zn1iii | 121.02 (5) |
| O3—Zn1—O2v | 84.54 (4) | As1—O2—Zn1viii | 126.04 (5) |
| O3i—Zn1—O2v | 163.24 (4) | Zn1iii—O2—Zn1viii | 101.35 (4) |
| O2ii—Zn1—O2v | 78.64 (4) | As1—O3—Zn1 | 141.36 (6) |
| Symmetry codes: (i) −x, y, −z+1/2; (ii) x−1/2, −y+1/2, z−1/2; (iii) −x+1/2, −y+1/2, −z+1; (iv) x−1/2, y−1/2, z; (v) −x+1/2, y−1/2, −z+1/2; (vi) −x+1/2, −y+1/2, −z; (vii) −x+1/2, y+1/2, −z+1/2; (viii) x+1/2, y+1/2, z. |
| Zn1 | As1 | Σ | |
| O1 | 1.10, 1.08 | 2.18 | |
| O2 | 0.36 (1×→, 2×↓), 0.29 (1×→, 2×↓) | 1.38 | 2.03 |
| O3 | 0.41 (1×→, 2×↓) | 1.45 | 1.86 |
| Σ | 2.12 | 5.01 |
Acknowledgements
TU Wien Bibliothek is acknowledged for financial support through its Open Access Funding Program.
References
 Aroyo, M. I., Perez-Mato, J. M., Capillas, C., Kroumova, E., Ivantchev, S., Madariaga, G., Kirov, A. & Wondratschek, H. (2006). Z. Kristallogr. 221, 15–27. Web of Science CrossRef CAS Google Scholar
Averbuch-Pouchot, M. T., Durif, A. & Bagieu-Beucher, M. (1983). Acta Cryst. C39, 25–26. CrossRef CAS Web of Science IUCr Journals Google Scholar
Bagieu-Beucher, M., Gondrand, M. & Perroux, M. (1976). J. Solid State Chem. 19, 353–357. CAS Google Scholar
Basso, R., Lucchetti, G., Zefiro, L. & Palenzona, A. (1996). Eur. J. Mineral. 8, 487–492. CrossRef CAS Google Scholar
Brown, I. D. (2002). The Chemical Bond in Inorganic Chemistry: The Bond Valence Model. Oxford University Press. Google Scholar
Brown, I. D. (2009). Chem. Rev. 109, 6858–6919. Web of Science CrossRef PubMed CAS Google Scholar
Dolomanov, O. V., Bourhis, L. J., Gildea, R. J., Howard, J. A. K. & Puschmann, H. (2009). J. Appl. Cryst. 42, 339–341. Web of Science CrossRef CAS IUCr Journals Google Scholar
Dowty, E. (2006). ATOMS. Shape Software, Kingsport, Tennessee, USA. Google Scholar
Durif, A. (1995). Crystal Chemistry of Condensed Phosphates pp. 167–168 New York and London: Plenum Press. Google Scholar
Flor, G. de la, Orobengoa, D., Tasci, E., Perez-Mato, J. M. & Aroyo, M. I. (2016). J. Appl. Cryst. 49, 653–664. Web of Science CrossRef IUCr Journals Google Scholar
Folkers-Karlsson, L. C., Celani, P., Hahn, F. & Richter, J. (2026). Z. Kristallogr. 241, 41–51. CAS Google Scholar
Frerichs, D., Park, C. H. & Müller-Buschbaum, Hk. (1996a). Z. Naturforsch. Teil B 51, 233–239. CrossRef CAS Google Scholar
Frerichs, D., Park, C. H. & Müller-Buschbaum, Hk. (1996b). Z. Naturforsch. Teil B 51, 333–337. CrossRef CAS Google Scholar
Gagné, O. C. & Hawthorne, F. C. (2018). Acta Cryst. B74, 63–78. Web of Science CrossRef IUCr Journals Google Scholar
Gagné, O. C. & Hawthorne, F. C. (2020). IUCrJ 7, 581–629. Web of Science CrossRef PubMed IUCr Journals Google Scholar
Gake, T., Hirai, D. & Hirose, S. (2025). Phys. Rev. Mater. 9, 113803. CrossRef Google Scholar
Griesemer, S. D., Baldassarri, B., Zhu, R., Shen, J., Pal, K., Park, C. W. & Wolverton, C. (2025). Sci. Adv. 11, eadq1431. CrossRef PubMed Google Scholar
Hill, R. J. (1987). J. Solid State Chem. 71, 12–18. CrossRef CAS Google Scholar
Ilinca, G. (2022). Minerals 12, 924. Web of Science CrossRef Google Scholar
Kasenov, B. K., Mustafin, E. S. & Sarkulov, Zh. Kh. (1996). Russ. J. Inorg. Chem. 41, 839–840. Google Scholar
Keller, P., Hess, H. & Riffel, H. (1980). Neues Jahrb. Mineral. Abh. 138, 316–332. CAS Google Scholar
Koziskova, J., Hahn, F., Richter, J. & Kožíšek, J. (2016). Acta Chim. Slovaca 9, 136–140. CrossRef CAS Google Scholar
Losilla, E. R., Aranda, M. A. G., Ramirez, F. J. & Bruque, S. (1995). J. Phys. Chem. 99, 12975–12979. CrossRef CAS Web of Science Google Scholar
Mustafin, E. S. & Kasenov, B. K. (1994). Russ. J. Inorg. Chem. 39, 716. Google Scholar
Mustafin, E. S. & Kasenov, B. K. (1996). J. Eng. Phys. Thermophys. 68, 529–531. CrossRef Google Scholar
Nakua, A. M. & Greedan, J. E. (1995). J. Solid State Chem. 118, 402–411. CrossRef CAS Web of Science Google Scholar
Orosel, D. & Jansen, M. (2006). Z. Anorg. Allg. Chem. 632, 1131–1133. Web of Science CrossRef CAS Google Scholar
Salinas-Sanchez, A., Garcia-Muñoz, J. L., Rodriguez-Carvajal, J., Saez-Puche, R. & Martinez, J. L. (1992). J. Solid State Chem. 100, 201–211. CAS Google Scholar
Sheldrick, G. M. (2015). Acta Cryst. C71, 3–8. Web of Science CrossRef IUCr Journals Google Scholar
Weil, M. (2000). Z. Naturforsch. Teil B 55, 699–706. CrossRef CAS Google Scholar
Weil, M. (2001). Acta Cryst. E57, i22–i23. Web of Science CrossRef IUCr Journals Google Scholar
Weil, M. (2004). Acta Cryst. C60, i20–i22. CrossRef CAS IUCr Journals Google Scholar
Weil, M. (2016). Cryst. Growth Des. 16, 908–921. Web of Science CrossRef CAS Google Scholar
Weil, M., Đorđević, T., Lengauer, C. L. & Kolitsch, U. (2009). Solid State Sci. 11, 2111–2117. Web of Science CrossRef CAS Google Scholar
Weil, M. & Stöger, B. (2010). Acta Cryst. B66, 603–614. Web of Science CrossRef ICSD CAS IUCr Journals Google Scholar
Westrip, S. P. (2010). J. Appl. Cryst. 43, 920–925. Web of Science CrossRef CAS IUCr Journals Google Scholar
Zagorac, D., Müller, H., Ruehl, S., Zagorac, J. & Rehme, S. (2019). J. Appl. Cryst. 52, 918–925. Web of Science CrossRef CAS IUCr Journals Google Scholar
This is an open-access article distributed under the terms of the Creative Commons Attribution (CC-BY) Licence, which permits unrestricted use, distribution, and reproduction in any medium, provided the original authors and source are cited.
| CRYSTALLOGRAPHIC COMMUNICATIONS |