Acta Crystallographica
Section E CRYSTALLOGRAPHIC COMMUNICATIONS


 menu


research communications
| CRYSTALLOGRAPHIC COMMUNICATIONS |
 access
Crystal structure and Hirshfeld surface analysis of 1,2-bis[5-(4-fluorophenyl)-1,3-oxazol-2-yl]benzene

aInstitute of Functional Materials Chemistry, SSI "Institute for Single Crystals" of National Academy of Sciences of Ukraine, Nauki Ave 60, Kharkiv 61001, Ukraine, bV. I. Vernadskii Institute of General and Inorganic Chemistry of, National Academy of Sciences of Ukraine, Prospect Palladina 32/34, Kyiv 03680, Ukraine, and cV. N. Karazin Kharkiv National University, 4 Svobody Sq., Kharkiv 61022, Ukraine
*Correspondence e-mail: [email protected]
The title compound, C24H14F2N2O2, was studied by single-crystal and powder X-ray diffraction methods. The title compound crystallizes in the monoclinic space group P21/c with one molecule in the asymmetric unit. The two chemically equivalent substituents adopt different orientations relative to the central benzene ring. The dihedral angles between the central benzene and oxazole rings are 2.4 (2) and 57.6 (2)°, while those between the oxazole and terminal 4-fluorophenyl rings are 13.3 (2) and 2.5 (2)°, respectively. In the crystal, inversion-related molecules form dimers through π–π stacking between oxazole rings [centroid-to-centroid distance = 3.587 (3) Å] and C—H⋯N hydrogen bonds. The dimers are further connected by C—H⋯N interactions into layers parallel to (100). Hirshfeld surface analysis indicates that C⋯H/H⋯C (30.2%), H⋯H (21.8%) and F⋯H/H⋯F (18.0%) contacts make the largest contributions to the crystal packing.
CCDC reference: 2586503
1. Chemical context
The title compound belongs to a series of sterically hindered ortho analogues of the well-known scintillator dye 1,4-bis(5-phenyl-1,3-oxazol-2-yl)benzene (POPOP; Ambats & Marsh, 1965 ). The approximately planar molecular structure of POPOP promotes extended π-conjugation and efficient fluorescence, accounting for its use in plastic and liquid scintillators (Ott et al., 1957). In ortho-POPOP, however, the two oxazolyl moieties occupy adjacent positions on the central benzene ring. Steric repulsion between these substituents results in pronounced molecular non-planarity both in the crystal and in solution (Doroshenko et al., 1994, 1997, 2000a, 2002a,b). The resulting decrease in π-conjugation produces a hypsochromic shift of the absorption spectrum (Doroshenko, 2002a). Partial flattening in the electronically excited state restores conjugation and gives rise to bathochromically shifted fluorescence and a large Stokes shift (Doroshenko et al., 1996, 2000b; Doroshenko, 1999; Iliashenko et al., 2011).
A characteristic feature of crystalline ortho-POPOP derivatives is that the two chemically equivalent oxazolyl substituents may adopt considerably different orientations relative to the central benzene ring (Doroshenko et al., 1994). The molecular conformation can be described in terms of a relatively planar three-ring benzene–oxazole–phenyl fragment and a two-ring oxazole–phenyl fragment that is strongly inclined to the central benzene ring. A closely related compound is the monofluoro-substituted derivative 5-(4-uflorophenyl)-2-[2-(5-phenyl-1,3-oxazol-2-yl)phenyl]-1,3-oxazole (2; Ilyashenko et al., 2010). In its crystal structure, the fluorine atom is disordered between the two terminal phenyl rings, with refined occupancies of 0.627 (3) and 0.373 (3).
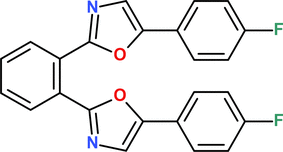
The title compound, 1,2-bis(5-(4-fluorophenyl)-1,3-oxazol-2-yl)benzene (1), is the corresponding difluoro-substituted ortho-POPOP derivative, in which both terminal phenyl rings bear a para-fluoro substituent. The present study reports its molecular and crystal structure and Hirshfeld surface analysis.
2. Structural commentary
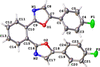
The molecular structure of the title compond 1 is shown in Fig. 1. The asymmetric unit contains one crystallographically independent molecule (Z′ = 1). The molecule consists of a central benzene ring bearing two 5-(4-fluorophenyl)-1,3-oxazol-2-yl moieties at adjacent ortho-positions.
|
Figure 1
The molecular structure of 1, showing the atom labelling and displacement ellipsoids drawn at the 50% probability level. |
The dihedral angles between the least-squares plane of the central benzene ring (C10–C15) and those of the oxazole rings O1/N1/C7–C9 and O2/N2/C16–C18 are 2.4 (2) and 57.6 (2)°, respectively. The corresponding dihedral angles between the oxazole rings and the terminal 4-fluorophenyl rings C1–C6 and C19–C24 are 13.4 (2) and 2.5 (2)°, respectively. Consequently, one moiety forms an approximately planar benzene–oxazole–4-fluorophenyl fragment, with only a slight rotation of the terminal 4-fluorophenyl ring. In the second moiety, the oxazole and 4-fluorophenyl rings are essentially coplanar, whereas their mean plane is strongly inclined to that of the central benzene ring. Thus, although the two substituents are chemically equivalent, they adopt different orientations relative to the central benzene ring in the crystal. Similar differences in the orientations of chemically equivalent substituents have been observed in other crystalline ortho-POPOP derivatives (Doroshenko et al., 1994, 1997, 2000a, 2002b).
3. Supramolecular features
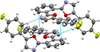
In the crystal, inversion-related molecules form dimers consolidated π–π stacking interactions between the O1/N1/C7–C9 oxazole rings (Fig. 2). The centroid-to-centroid distance Cg1⋯Cg1iv is 3.587 (3) Å, with a perpendicular distance of 3.456 Å and a shift of 0.962 Å [Cg1 is the centroid of the O1/N1/C7–C9 ring; symmetry code: (iv) 1 − x, 1 − y, 1 – z]. The dimers are additionally consolidated by pairs of C24—H24⋯N1iv hydrogen bonds (Table 1).
|
 ; (ii)
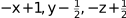 ; (iii)
 ; (iv)
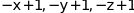 .
|
Figure 2
A centrosymmetric dimer of 1 formed by π–π stacking and C—H⋯N hydrogen bonds. The intermolecular interactions are shown as dashed lines. [Symmetry code: (iv) 1 − x, 1 − y, 1 − z.] |
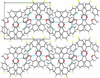
The inversion-related dimers are connected by C8—H8⋯N2ii hydrogen bonds, forming chains extending parallel to [010]. Together, the C—H⋯N hydrogen bonds and π–π interactions generate layers parallel to (100), as shown in Fig. 3. The terminal fluorine atoms are directed towards the interlayer region. Weak C5—H5⋯F2i contacts are observed between adjacent layers along the a-axis direction (Table 1) and may contribute to the consolidation of the crystal packing (Thalladi et al., 1998). A further weak C23—H23⋯F1iii contact occurs within the layers (Table 1).
|
Figure 3
Crystal packing of 1 showing the C—H⋯N hydrogen-bonded layers parallel to (100). |
4. Powder X-ray diffraction
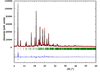
The powder diffraction pattern of 1 was recorded using a Rigaku SmartLab powder diffractometer at room temperature (Cu Kα radiation, Bragg–Brentano geometry, θ–θ scanning, Ni filter, 3° < 2θ < 60°, Δ2θ = 0.01°). The Rietveld refinement of the obtained pattern (Fig. 4) was carried out using the FullProf and WinPLOTR programs (Rodríguez-Carvajal & Roisnel, 1998), with data from the external NIST SRM 640 standard used for the calculation of the instrumental profile function and the single-crystal structure used as the structural model for refinement. A minor preferred orientation effect along the [110] direction was revealed during the refinement and taken into account. The crystallite size obtained from the refinement was approximately 99 nm, indicating relatively large crystallites and no pronounced nanocrystalline character of the sample. The main results of the Rietveld refinement are summarized in Table 2. The experimental powder X-ray diffraction pattern is in good agreement with the theoretical pattern calculated from the single-crystal structure data.
|
||||||||||||||||||||||||||||||||||
|
Figure 4
Final Rietveld plot of compound 1. Observed data points are indicated by red dots, the best-fit profile (upper trace) and the difference pattern (lower trace) are shown as solid lines. The vertical bars correspond to the positions of the Bragg peaks. |
5. Hirshfeld surface analysis and fingerprint plots
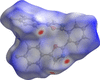
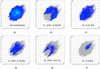
A Hirshfeld surface analysis was performed using CrystalExplorer21 (Spackman et al., 2021) to examine the intermolecular contacts in the crystal of 1. The Hirshfeld surface mapped over dnorm (Spackman & Jayatilaka, 2009) is shown in Fig. 5, while the corresponding two-dimensional fingerprint plots (McKinnon et al., 2007) are presented in Fig. 6. The red spots on the dnorm surface correspond to the C—H⋯N contacts described above.
|
Figure 5
Three-dimensional Hirshfeld surface of compound 1 mapped over dnorm. |
|
Figure 6
Two-dimensional fingerprint plots for compound 1 showing (a) all intermolecular contacts and the plots delineated into contributions from (b) C⋯H/H⋯C, (c) H⋯H, (d) F⋯H/H⋯F, (e) N⋯H/H⋯N, (f) C⋯C contacts. The quantities de and di represent the distances from a point on the Hirshfeld surface to the nearest atom outside and inside the surface, respectively. |
The decomposed fingerprint plots show that C⋯H/H⋯C contacts make the largest contribution to the Hirshfeld surface, accounting for 30.2%, followed by H⋯H contacts (21.8%) and F⋯H/H⋯F contacts (18.0%). The relatively high contribution of F⋯H/H⋯F contacts reflects the frequent close approaches between the fluorine atoms and hydrogen atoms of neighbouring molecules. These include the weak interlayer C5—H5⋯F2i and intralayer C23—H23⋯F1iii contacts described above. Thus, contacts involving fluorine atoms contribute to the molecular association both within and between the layers.
The N⋯H/H⋯N contacts account for 8.1% of the Hirshfeld surface and are consistent with the C—H⋯N interactions described in the Supramolecular features section. The C⋯C contacts contribute 5.4% to the Hirshfeld surface and support the presence of π–π stacking interactions between inversion-related oxazole rings.
Smaller contributions arise from O⋯H/H⋯O and O⋯N/N⋯O contacts, each accounting for 2.9%, as well as from N⋯C/C⋯N (1.5%), F⋯F (1.4%), O⋯C/C⋯O (1.0%) and F⋯C/C⋯F (0.8%) contacts. These minor contributions represent other weak contacts present in the crystal packing.
6. Database survey
An expanded substructure search of the Cambridge Structural Database (CSD, Version 6.01, updated February 2026; Groom et al., 2016), based on the ortho-disubstituted central benzene core and related five-membered O,N-heterocycles, gave ten entries. Three hits corresponding to bis(imidazole) derivatives (GUVNET, GUVNET01 and SODQEJ) were excluded because they do not contain oxazole or oxadiazole rings. The remaining seven entries correspond to six unique related crystal structures, since PODZEN and PODZEN10 describe the same crystallographic model.
The closest bis(1,3-oxazole) analogues are the unsubstituted ortho-POPOP molecule (PODZEN/PODZEN10; Doroshenko et al., 1994) and its monofluoro derivative (SUZVEP; Ilyashenko et al., 2010). Three unsymmetrical analogues containing one 1,3-oxazole and one 1,3,4-oxadiazole ring were also found: NODYOU (Doroshenko et al., 1997), EDEFIC (Doroshenko et al., 2000a) and HUGJUO (Doroshenko et al., 2002b). The more distant bis(1,3,4-oxadiazole) analogue SEQREN was reported by Yang et al. (2010).
All these structures adopt non-planar molecular geometries in which the two substituents have different orientations relative to the central benzene ring. In PODZEN/PODZEN10, the dihedral angles between the plane of the central benzene ring and the planes of the two oxazole rings are 10.3 and 66.9°, respectively. The corresponding published values for SUZVEP are 10.7 and 64.1° (Ilyashenko et al., 2010). In the title compound, these angles are 2.37 and 57.60°, indicating somewhat smaller twisting of both oxazole rings relative to the central benzene ring, although the difference between the orientations of the two oxazole moieties, characteristic of the ortho-POPOP framework, is retained.
7. Synthesis and crystallization
The title compound 1 was synthesized by the reported procedure (Doroshenko et al., 1994) using phthaloyl dichloride and 4-F-substituted ω-amino-acetophenone hydrochloride (Ilyashenko et al., 2010). A solution of phthaloyl dichloride (0.025 mol, ∼3.6 mL) in 20 mL of benzene was mixed with 9.5 g (0.05 mol) 4-F-ω-aminoacetophenone hydrochloride in 30 mL of water. The reaction mixture was basified dropwise with concentrated aqueous sodium carbonate with continuous stirring to keep it weakly alkaline and the mixture was stirred for 1 h. Then the resulting precipitate was filtered off, washed with water and air-dried at room temperature. The obtained intermediate was dissolved in 80 ml of concentrated sulfuric acid, stirred 30 min at room temperature, and then for an additional 30 min at 328 K. The reaction mixture was cooled, poured onto ice, the product was filtered off, thoroughly washed with water until the washings were neutral, air-dried and finally recrystallized from ethanol. Yield: 5.9 g (58%); colorless crystals.
Crystals suitable for X-ray diffraction analysis were obtained by slow evaporation of a concentrated solution of 1 in propan-2-ol.
8. Refinement
Crystal data, data collection and structure refinement details are summarized in Table 3. The H atoms were placed in calculated positions (C—H = 0.93 Å) and refined using a riding model with Uiso(H) = 1.2Ueq of the carrier atom.
|
Supporting information
CCDC reference: 2586503
Crystal structure: contains datablock I. DOI: https://doi.org/10.1107/S2056989026009394/ex2105sup1.cif
Structure factors: contains datablock I. DOI: https://doi.org/10.1107/S2056989026009394/ex2105Isup2.hkl
Supporting information file. DOI: https://doi.org/10.1107/S2056989026009394/ex2105Isup3.cml
| C24H14F2N2O2 | F(000) = 824 |
| Mr = 400.37 | Dx = 1.396 Mg m−3 |
| Monoclinic, P21/c | Mo Kα radiation, λ = 0.71073 Å |
| a = 13.767 (8) Å | Cell parameters from 633 reflections |
| b = 16.914 (11) Å | θ = 2.4–22.8° |
| c = 8.447 (5) Å | µ = 0.10 mm−1 |
| β = 104.423 (13)° | T = 296 K |
| V = 1904.9 (19) Å3 | Block, colourless |
| Z = 4 | 0.3 × 0.2 × 0.1 mm |
| Bruker APEXII CCD diffractometer | 1446 reflections with I > 2σ(I) |
| φ and ω scans | Rint = 0.110 |
| Absorption correction: multi-scan (SADABS; Krause et al., 2015) | θmax = 26.0°, θmin = 1.5° |
| Tmin = 0.617, Tmax = 0.746 | h = −16→16 |
| 10872 measured reflections | k = −20→20 |
| 3740 independent reflections | l = −8→10 |
| Refinement on F2 | Hydrogen site location: inferred from neighbouring sites |
| Least-squares matrix: full | H-atom parameters constrained |
| R[F2 > 2σ(F2)] = 0.076 | w = 1/[σ2(Fo2) + (0.045P)2] where P = (Fo2 + 2Fc2)/3 |
| wR(F2) = 0.151 | (Δ/σ)max < 0.001 |
| S = 0.91 | Δρmax = 0.20 e Å−3 |
| 3740 reflections | Δρmin = −0.19 e Å−3 |
| 272 parameters | Extinction correction: SHELXL2019/3 (Sheldrick, 2015b), Fc*=kFc[1+0.001xFc2λ3/sin(2θ)]-1/4 |
| 0 restraints | Extinction coefficient: 0.0043 (8) |
Geometry. All esds (except the esd in the dihedral angle between two l.s. planes) are estimated using the full covariance matrix. The cell esds are taken into account individually in the estimation of esds in distances, angles and torsion angles; correlations between esds in cell parameters are only used when they are defined by crystal symmetry. An approximate (isotropic) treatment of cell esds is used for estimating esds involving l.s. planes. |
Refinement. Using Olex2 (Dolomanov et al., 2009), the structure was solved with the SHELXT (Sheldrick, 2015a) structure solution program using Intrinsic Phasing and refined with the SHELXL (Sheldrick, 2015b) refinement package. Full-matrix least squares refinement against F2 in anisotropic approximation was used for non-hydrogen atoms. |
| x | y | z | Uiso*/Ueq | ||
| F1 | 0.0504 (2) | 0.4923 (2) | −0.1473 (3) | 0.1238 (14) | |
| F2 | −0.06623 (16) | 0.61801 (17) | 0.4548 (3) | 0.0916 (10) | |
| O1 | 0.47053 (18) | 0.58652 (15) | 0.3229 (3) | 0.0414 (7) | |
| O2 | 0.39878 (17) | 0.68355 (14) | 0.5272 (3) | 0.0409 (7) | |
| N1 | 0.6035 (2) | 0.5080 (2) | 0.3606 (4) | 0.0538 (10) | |
| N2 | 0.4544 (2) | 0.7954 (2) | 0.4465 (4) | 0.0585 (10) | |
| C1 | 0.3414 (3) | 0.5178 (3) | 0.1210 (5) | 0.0474 (11) | |
| C2 | 0.2666 (3) | 0.5654 (3) | 0.1507 (5) | 0.0583 (13) | |
| H2 | 0.282616 | 0.603993 | 0.231339 | 0.070* | |
| C3 | 0.1670 (3) | 0.5564 (3) | 0.0611 (6) | 0.0762 (16) | |
| H3 | 0.115874 | 0.587502 | 0.082073 | 0.091* | |
| C4 | 0.1481 (4) | 0.5002 (4) | −0.0581 (6) | 0.0791 (17) | |
| C5 | 0.2189 (4) | 0.4515 (3) | −0.0906 (6) | 0.0819 (18) | |
| H5 | 0.202142 | 0.413168 | −0.171779 | 0.098* | |
| C6 | 0.3169 (3) | 0.4603 (3) | 0.0002 (5) | 0.0675 (14) | |
| H6 | 0.366723 | 0.427564 | −0.019925 | 0.081* | |
| C7 | 0.4445 (3) | 0.5227 (2) | 0.2193 (5) | 0.0399 (10) | |
| C8 | 0.5244 (3) | 0.4759 (2) | 0.2423 (5) | 0.0551 (12) | |
| H8 | 0.526946 | 0.428583 | 0.187402 | 0.066* | |
| C9 | 0.5673 (3) | 0.5719 (2) | 0.4056 (5) | 0.0386 (10) | |
| C10 | 0.6219 (3) | 0.6282 (2) | 0.5302 (4) | 0.0360 (10) | |
| C11 | 0.7223 (3) | 0.6099 (2) | 0.6003 (5) | 0.0517 (12) | |
| H11 | 0.749581 | 0.564028 | 0.568470 | 0.062* | |
| C12 | 0.7819 (3) | 0.6586 (3) | 0.7162 (5) | 0.0623 (14) | |
| H12 | 0.848343 | 0.645215 | 0.763227 | 0.075* | |
| C13 | 0.7421 (3) | 0.7271 (3) | 0.7613 (5) | 0.0643 (14) | |
| H13 | 0.782182 | 0.760607 | 0.837773 | 0.077* | |
| C14 | 0.6433 (3) | 0.7465 (3) | 0.6938 (5) | 0.0555 (12) | |
| H14 | 0.617286 | 0.792681 | 0.726634 | 0.067* | |
| C15 | 0.5814 (3) | 0.6980 (2) | 0.5770 (5) | 0.0392 (10) | |
| C16 | 0.4786 (3) | 0.7270 (2) | 0.5112 (5) | 0.0400 (10) | |
| C17 | 0.3507 (3) | 0.7967 (3) | 0.4137 (5) | 0.0591 (13) | |
| H17 | 0.310674 | 0.838583 | 0.364357 | 0.071* | |
| C18 | 0.3158 (3) | 0.7298 (2) | 0.4625 (5) | 0.0438 (11) | |
| C19 | 0.2165 (3) | 0.6990 (2) | 0.4603 (5) | 0.0447 (11) | |
| C20 | 0.1319 (3) | 0.7432 (3) | 0.3893 (6) | 0.0688 (14) | |
| H20 | 0.139112 | 0.792099 | 0.343075 | 0.083* | |
| C21 | 0.0372 (3) | 0.7154 (3) | 0.3868 (6) | 0.0789 (16) | |
| H21 | −0.019404 | 0.744990 | 0.339041 | 0.095* | |
| C22 | 0.0282 (3) | 0.6444 (3) | 0.4549 (6) | 0.0614 (13) | |
| C23 | 0.1077 (3) | 0.5980 (3) | 0.5230 (5) | 0.0606 (13) | |
| H23 | 0.098947 | 0.549031 | 0.567457 | 0.073* | |
| C24 | 0.2030 (3) | 0.6262 (3) | 0.5245 (5) | 0.0501 (11) | |
| H24 | 0.258774 | 0.595131 | 0.569852 | 0.060* |
| U11 | U22 | U33 | U12 | U13 | U23 | |
| F1 | 0.0618 (19) | 0.213 (4) | 0.087 (2) | −0.037 (2) | 0.0004 (17) | −0.033 (2) |
| F2 | 0.0395 (15) | 0.121 (3) | 0.114 (2) | −0.0090 (16) | 0.0200 (15) | −0.004 (2) |
| O1 | 0.0399 (16) | 0.0409 (17) | 0.0429 (17) | −0.0019 (14) | 0.0093 (13) | −0.0034 (14) |
| O2 | 0.0350 (15) | 0.0380 (16) | 0.0513 (18) | 0.0020 (14) | 0.0137 (13) | 0.0023 (14) |
| N1 | 0.048 (2) | 0.048 (2) | 0.065 (3) | 0.0102 (19) | 0.014 (2) | −0.004 (2) |
| N2 | 0.050 (2) | 0.047 (2) | 0.082 (3) | −0.0011 (19) | 0.021 (2) | 0.007 (2) |
| C1 | 0.045 (3) | 0.055 (3) | 0.045 (3) | −0.008 (2) | 0.016 (2) | −0.006 (2) |
| C2 | 0.046 (3) | 0.080 (4) | 0.049 (3) | −0.009 (3) | 0.013 (2) | −0.012 (3) |
| C3 | 0.050 (3) | 0.106 (5) | 0.072 (4) | −0.003 (3) | 0.013 (3) | −0.006 (3) |
| C4 | 0.052 (4) | 0.128 (6) | 0.052 (3) | −0.031 (3) | 0.005 (3) | −0.012 (3) |
| C5 | 0.066 (4) | 0.121 (5) | 0.058 (4) | −0.026 (4) | 0.016 (3) | −0.032 (3) |
| C6 | 0.063 (3) | 0.080 (4) | 0.061 (3) | −0.014 (3) | 0.017 (3) | −0.016 (3) |
| C7 | 0.046 (3) | 0.038 (3) | 0.036 (3) | −0.008 (2) | 0.011 (2) | −0.005 (2) |
| C8 | 0.059 (3) | 0.042 (3) | 0.066 (3) | 0.001 (3) | 0.017 (3) | −0.014 (2) |
| C9 | 0.037 (2) | 0.040 (3) | 0.039 (3) | 0.001 (2) | 0.012 (2) | 0.010 (2) |
| C10 | 0.033 (2) | 0.040 (2) | 0.036 (2) | −0.002 (2) | 0.0108 (19) | 0.006 (2) |
| C11 | 0.037 (2) | 0.050 (3) | 0.069 (3) | 0.002 (2) | 0.015 (2) | 0.008 (3) |
| C12 | 0.034 (3) | 0.083 (4) | 0.064 (3) | −0.012 (3) | 0.002 (2) | 0.008 (3) |
| C13 | 0.047 (3) | 0.076 (4) | 0.067 (3) | −0.018 (3) | 0.009 (2) | −0.020 (3) |
| C14 | 0.052 (3) | 0.058 (3) | 0.057 (3) | −0.007 (2) | 0.014 (2) | −0.011 (2) |
| C15 | 0.036 (2) | 0.047 (3) | 0.036 (2) | −0.003 (2) | 0.012 (2) | 0.003 (2) |
| C16 | 0.043 (3) | 0.034 (3) | 0.046 (3) | −0.008 (2) | 0.016 (2) | −0.007 (2) |
| C17 | 0.053 (3) | 0.049 (3) | 0.076 (3) | 0.013 (2) | 0.018 (3) | 0.014 (3) |
| C18 | 0.043 (3) | 0.044 (3) | 0.044 (3) | 0.002 (2) | 0.010 (2) | 0.005 (2) |
| C19 | 0.043 (3) | 0.046 (3) | 0.045 (3) | 0.008 (2) | 0.012 (2) | 0.002 (2) |
| C20 | 0.048 (3) | 0.062 (3) | 0.096 (4) | 0.011 (3) | 0.017 (3) | 0.026 (3) |
| C21 | 0.037 (3) | 0.087 (4) | 0.110 (4) | 0.019 (3) | 0.014 (3) | 0.017 (4) |
| C22 | 0.036 (3) | 0.078 (4) | 0.071 (4) | −0.002 (3) | 0.013 (2) | −0.002 (3) |
| C23 | 0.054 (3) | 0.063 (3) | 0.066 (3) | −0.007 (3) | 0.016 (3) | 0.004 (3) |
| C24 | 0.032 (2) | 0.058 (3) | 0.057 (3) | 0.008 (2) | 0.005 (2) | −0.001 (2) |
| F1—C4 | 1.374 (5) | C10—C11 | 1.396 (4) |
| F2—C22 | 1.375 (5) | C10—C15 | 1.404 (5) |
| O1—C7 | 1.379 (4) | C11—H11 | 0.9300 |
| O1—C9 | 1.365 (4) | C11—C12 | 1.382 (5) |
| O2—C16 | 1.356 (4) | C12—H12 | 0.9300 |
| O2—C18 | 1.380 (4) | C12—C13 | 1.376 (6) |
| N1—C8 | 1.392 (5) | C13—H13 | 0.9300 |
| N1—C9 | 1.286 (4) | C13—C14 | 1.377 (5) |
| N2—C16 | 1.288 (5) | C14—H14 | 0.9300 |
| N2—C17 | 1.386 (5) | C14—C15 | 1.397 (5) |
| C1—C2 | 1.378 (5) | C15—C16 | 1.470 (5) |
| C1—C6 | 1.388 (5) | C17—H17 | 0.9300 |
| C1—C7 | 1.458 (5) | C17—C18 | 1.334 (5) |
| C2—H2 | 0.9300 | C18—C19 | 1.459 (5) |
| C2—C3 | 1.401 (5) | C19—C20 | 1.388 (5) |
| C3—H3 | 0.9300 | C19—C24 | 1.376 (5) |
| C3—C4 | 1.362 (6) | C20—H20 | 0.9300 |
| C4—C5 | 1.356 (6) | C20—C21 | 1.382 (6) |
| C5—H5 | 0.9300 | C21—H21 | 0.9300 |
| C5—C6 | 1.383 (5) | C21—C22 | 1.349 (6) |
| C6—H6 | 0.9300 | C22—C23 | 1.352 (5) |
| C7—C8 | 1.329 (5) | C23—H23 | 0.9300 |
| C8—H8 | 0.9300 | C23—C24 | 1.394 (5) |
| C9—C10 | 1.479 (5) | C24—H24 | 0.9300 |
| C9—O1—C7 | 104.4 (3) | C13—C12—C11 | 119.4 (4) |
| C16—O2—C18 | 105.4 (3) | C13—C12—H12 | 120.3 |
| C9—N1—C8 | 104.4 (3) | C12—C13—H13 | 119.8 |
| C16—N2—C17 | 104.0 (3) | C12—C13—C14 | 120.4 (4) |
| C2—C1—C6 | 119.2 (4) | C14—C13—H13 | 119.8 |
| C2—C1—C7 | 121.9 (4) | C13—C14—H14 | 119.4 |
| C6—C1—C7 | 118.9 (4) | C13—C14—C15 | 121.2 (4) |
| C1—C2—H2 | 119.5 | C15—C14—H14 | 119.4 |
| C1—C2—C3 | 120.9 (4) | C10—C15—C16 | 125.9 (3) |
| C3—C2—H2 | 119.5 | C14—C15—C10 | 118.5 (4) |
| C2—C3—H3 | 121.4 | C14—C15—C16 | 115.6 (4) |
| C4—C3—C2 | 117.1 (5) | O2—C16—C15 | 120.6 (3) |
| C4—C3—H3 | 121.4 | N2—C16—O2 | 113.6 (3) |
| C3—C4—F1 | 117.0 (5) | N2—C16—C15 | 125.6 (4) |
| C5—C4—F1 | 119.0 (5) | N2—C17—H17 | 124.5 |
| C5—C4—C3 | 124.0 (5) | C18—C17—N2 | 111.0 (4) |
| C4—C5—H5 | 120.9 | C18—C17—H17 | 124.5 |
| C4—C5—C6 | 118.2 (5) | O2—C18—C19 | 118.9 (4) |
| C6—C5—H5 | 120.9 | C17—C18—O2 | 106.1 (4) |
| C1—C6—H6 | 119.7 | C17—C18—C19 | 135.0 (4) |
| C5—C6—C1 | 120.5 (5) | C20—C19—C18 | 119.7 (4) |
| C5—C6—H6 | 119.7 | C24—C19—C18 | 122.1 (4) |
| O1—C7—C1 | 117.9 (4) | C24—C19—C20 | 118.2 (4) |
| C8—C7—O1 | 107.4 (3) | C19—C20—H20 | 119.7 |
| C8—C7—C1 | 134.5 (4) | C21—C20—C19 | 120.6 (4) |
| N1—C8—H8 | 125.0 | C21—C20—H20 | 119.7 |
| C7—C8—N1 | 110.1 (4) | C20—C21—H21 | 120.6 |
| C7—C8—H8 | 125.0 | C22—C21—C20 | 118.9 (4) |
| O1—C9—C10 | 120.4 (3) | C22—C21—H21 | 120.6 |
| N1—C9—O1 | 113.7 (3) | C21—C22—F2 | 118.4 (4) |
| N1—C9—C10 | 125.9 (4) | C21—C22—C23 | 123.2 (4) |
| C11—C10—C9 | 115.6 (4) | C23—C22—F2 | 118.4 (5) |
| C11—C10—C15 | 119.2 (4) | C22—C23—H23 | 121.1 |
| C15—C10—C9 | 125.1 (3) | C22—C23—C24 | 117.8 (4) |
| C10—C11—H11 | 119.4 | C24—C23—H23 | 121.1 |
| C12—C11—C10 | 121.2 (4) | C19—C24—C23 | 121.4 (4) |
| C12—C11—H11 | 119.4 | C19—C24—H24 | 119.3 |
| C11—C12—H12 | 120.3 | C23—C24—H24 | 119.3 |
| F1—C4—C5—C6 | 179.9 (4) | C9—C10—C15—C14 | −178.3 (4) |
| F2—C22—C23—C24 | −179.0 (4) | C9—C10—C15—C16 | 0.1 (6) |
| O1—C7—C8—N1 | 0.0 (5) | C10—C11—C12—C13 | −1.1 (6) |
| O1—C9—C10—C11 | −176.3 (3) | C10—C15—C16—O2 | 61.0 (5) |
| O1—C9—C10—C15 | 1.6 (6) | C10—C15—C16—N2 | −124.5 (5) |
| O2—C18—C19—C20 | −177.2 (4) | C11—C10—C15—C14 | −0.5 (6) |
| O2—C18—C19—C24 | 1.8 (6) | C11—C10—C15—C16 | 177.9 (4) |
| N1—C9—C10—C11 | 1.7 (6) | C11—C12—C13—C14 | 1.1 (7) |
| N1—C9—C10—C15 | 179.6 (4) | C12—C13—C14—C15 | −0.8 (7) |
| N2—C17—C18—O2 | −0.5 (5) | C13—C14—C15—C10 | 0.5 (6) |
| N2—C17—C18—C19 | 179.7 (4) | C13—C14—C15—C16 | −178.1 (4) |
| C1—C2—C3—C4 | −1.5 (7) | C14—C15—C16—O2 | −120.5 (4) |
| C1—C7—C8—N1 | −175.8 (4) | C14—C15—C16—N2 | 54.0 (6) |
| C2—C1—C6—C5 | 0.7 (7) | C15—C10—C11—C12 | 0.8 (6) |
| C2—C1—C7—O1 | −12.4 (6) | C16—O2—C18—C17 | −0.3 (4) |
| C2—C1—C7—C8 | 163.0 (5) | C16—O2—C18—C19 | 179.5 (3) |
| C2—C3—C4—F1 | −179.1 (4) | C16—N2—C17—C18 | 1.2 (5) |
| C2—C3—C4—C5 | 2.2 (8) | C17—N2—C16—O2 | −1.4 (5) |
| C3—C4—C5—C6 | −1.4 (9) | C17—N2—C16—C15 | −176.2 (4) |
| C4—C5—C6—C1 | −0.1 (7) | C17—C18—C19—C20 | 2.4 (8) |
| C6—C1—C2—C3 | 0.1 (7) | C17—C18—C19—C24 | −178.5 (5) |
| C6—C1—C7—O1 | 171.3 (4) | C18—O2—C16—N2 | 1.1 (4) |
| C6—C1—C7—C8 | −13.3 (7) | C18—O2—C16—C15 | 176.2 (3) |
| C7—O1—C9—N1 | 1.9 (4) | C18—C19—C20—C21 | −179.6 (4) |
| C7—O1—C9—C10 | −179.9 (3) | C18—C19—C24—C23 | 179.2 (4) |
| C7—C1—C2—C3 | −176.2 (4) | C19—C20—C21—C22 | 0.2 (8) |
| C7—C1—C6—C5 | 177.1 (4) | C20—C19—C24—C23 | −1.7 (6) |
| C8—N1—C9—O1 | −1.9 (5) | C20—C21—C22—F2 | 178.6 (4) |
| C8—N1—C9—C10 | −179.9 (4) | C20—C21—C22—C23 | −1.4 (8) |
| C9—O1—C7—C1 | 175.5 (3) | C21—C22—C23—C24 | 1.0 (7) |
| C9—O1—C7—C8 | −1.0 (4) | C22—C23—C24—C19 | 0.6 (6) |
| C9—N1—C8—C7 | 1.1 (5) | C24—C19—C20—C21 | 1.3 (7) |
| C9—C10—C11—C12 | 178.8 (4) |
| D—H···A | D—H | H···A | D···A | D—H···A |
| C5—H5···F2i | 0.93 | 2.70 | 3.470 (5) | 141 |
| C8—H8···N2ii | 0.93 | 2.56 | 3.489 (6) | 176 |
| C23—H23···F1iii | 0.93 | 2.82 | 3.559 (5) | 137 |
| C24—H24···N1iv | 0.93 | 2.54 | 3.449 (5) | 168 |
| Symmetry codes: (i) −x, −y+1, −z; (ii) −x+1, y−1/2, −z+1/2; (iii) x, y, z+1; (iv) −x+1, −y+1, −z+1. |
| Crystal system, space group | Monoclinic, P21/c |
| a (Å) | 13.7470 (5) |
| b (Å) | 16.9219 (4) |
| c (Å) | 8.4473 (1) |
| β (°) | 104.432 (2) |
| V (Å3) | 1903.04 (8) |
| Dx, Mg m-3 | 1.397 |
| Apparent crystallite size (nm) | 99 |
| Refinement | |
| Rp | 0.0590 |
| Rwp | 0.0942 |
| Rexp | 0.0147 |
| Rb | 0.0592 |
| Rf | 0.0561 |
Funding information
Funding for this research was provided by: National Academy of Science of Ukraine (grant No. 0125U000623 to V. V. Dyakonenko, A. M. Shaposhnyk, A. V. Kyrychenko).
References
 Ambats, I. & Marsh, R. E. (1965). Acta Cryst. 19, 942–948.
CrossRef
CAS
IUCr Journals
Web of Science
Google Scholar
Bruker (2008). APEX2 and SAINT. Bruker AXS Inc., Madison, Wisconsin, USA.
Google Scholar
Dolomanov, O. V., Bourhis, L. J., Gildea, R. J., Howard, J. A. K. & Puschmann, H. (2009). J. Appl. Cryst. 42, 339–341.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Doroshenko, A. (1999). Chem. Phys. Rep. 18, 873-879.
Google Scholar
Doroshenko, A. O. (2002a). Theor. Exp. Chem. 38, 135–155.
CrossRef
CAS
Google Scholar
Doroshenko, A. O., Baumer, V. N., Kirichenko, A. V., Shershukov, V. M. & Tolmachev, A. V. (1997). Chem. Heterocycl. Compd. 33, 1341–1349.
CrossRef
CAS
Google Scholar
Doroshenko, A. O., Baumer, V. N., Verezubova, A. A. & Ptyagina, L. M. (2002b). J. Mol. Struct. 609, 29–37.
Web of Science
CrossRef
CAS
Google Scholar
Doroshenko, A. O., Kirichenko, A. V., Mitina, V. G. & Ponomaryov, O. A. (1996). J. Photochem. Photobiol. Chem. 94, 15–26.
CrossRef
CAS
Google Scholar
Doroshenko, A. O., Kyrychenko, A. V., Baumer, V. N., Verezubova, A. A. & Ptyagina, L. M. (2000a). J. Mol. Struct. 524, 289–296.
CrossRef
CAS
Google Scholar
Doroshenko, A. O., Kyrychenko, A. V. & Waluk, J. (2000b). J. Fluoresc. 10, 41–48.
CrossRef
CAS
Google Scholar
Doroshenko, A. O., Patsenker, L. D., Baumer, V. N., Chepeleva, L. V., Van'Kevich, A. V., Kirichenko, A. V., Yarmolenko, S. N., Scherschukov, V. M., Mitina, V. G. & Ponomaryov, O. A. (1994). Mol. Eng. 3, 353–363.
CrossRef
CAS
Google Scholar
Groom, C. R., Bruno, I. J., Lightfoot, M. P. & Ward, S. C. (2016). Acta Cryst. B72, 171–179.
Web of Science
CrossRef
IUCr Journals
Google Scholar
Iliashenko, R., Zozulia, O. & Doroshenko, A. (2011). Centr. Eur. J. Chem. 9, 962-971.
CAS
Google Scholar
Ilyashenko, R., Wera, M., Błażejowski, J. & Doroshenko, A. (2010). Acta Cryst. E66, o2379–o2380.
CrossRef
IUCr Journals
Google Scholar
Krause, L., Herbst-Irmer, R., Sheldrick, G. M. & Stalke, D. (2015). J. Appl. Cryst. 48, 3–10.
Web of Science
CSD
CrossRef
ICSD
CAS
IUCr Journals
Google Scholar
McKinnon, J. J., Jayatilaka, D. & Spackman, M. A. (2007). Chem. Commun. pp. 3814–3816.
Web of Science
CrossRef
Google Scholar
Ott, D. G., Hayes, F. N., Hansbury, E. & Kerr, V. N. (1957). J. Am. Chem. Soc. 79, 5448–5454.
CrossRef
CAS
Google Scholar
Rodríguez-Carvajal, J. & Roisnel, T. (1998). IUCr Newsletter 20, 35–36.
Google Scholar
Sheldrick, G. M. (2015a). Acta Cryst. A71, 3–8.
Web of Science
CrossRef
IUCr Journals
Google Scholar
Sheldrick, G. M. (2015b). Acta Cryst. C71, 3–8.
Web of Science
CrossRef
IUCr Journals
Google Scholar
Spackman, M. A. & Jayatilaka, D. (2009). CrystEngComm 11, 19–32.
Web of Science
CrossRef
CAS
Google Scholar
Spackman, P. R., Turner, M. J., McKinnon, J. J., Wolff, S. K., Grimwood, D. J., Jayatilaka, D. & Spackman, M. A. (2021). J. Appl. Cryst. 54, 1006–1011.
Web of Science
CrossRef
CAS
IUCr Journals
Google Scholar
Thalladi, V. R., Weiss, H.-C., Bläser, D., Boese, R., Nangia, A. & Desiraju, G. R. (1998). J. Am. Chem. Soc. 120, 8702–8710.
Web of Science
CSD
CrossRef
CAS
Google Scholar
Yang, C.-C., Hsu, C.-J., Chou, P.-T., Cheng, H.-C., Su, Y. O. & Leung, M.-K. (2010). J. Phys. Chem. B 114, 756–768.
CrossRef
CAS
PubMed
Google Scholar
This is an open-access article distributed under the terms of the Creative Commons Attribution (CC-BY) Licence, which permits unrestricted use, distribution, and reproduction in any medium, provided the original authors and source are cited.
| CRYSTALLOGRAPHIC COMMUNICATIONS |