Acta Crystallographica
Section E CRYSTALLOGRAPHIC COMMUNICATIONS


 menu


research communications
| CRYSTALLOGRAPHIC COMMUNICATIONS |
 access
structure of the trigonal mixed-metal phosphate RbCoFe(PO4)2

aLaboratoire de Chimie Appliquée des Matériaux, Centre des Sciences des Matériaux, Faculty of Science, Mohammed V University in Rabat, Avenue Ibn Batouta, BP 1014, Rabat, Morocco, and bLaboratoire de Chimie des Matériaux Inorganiques, Faculté des Sciences, Département de Chimie, Université des Sciences et Techniques de Masuku, BP 943, Franceville, Gabon
*Correspondence e-mail: [email protected]
The title compound, rubidium cobalt iron bis(phosphate), was synthesized by the solid-state method under atmospheric conditions. This phosphate crystallizes with trigonal symmetry in P3m1 and all atoms lie on special crystallographic positions. In this structure, the CoIIO4 tetrahedra alternate with one of the two types of PO4 tetrahedra, sharing the vertices of their bases and extending along the a- and b-axis directions to form a first tetrahedral layer, lying parallel to the (001) plane. The same alternation is observed between the FeIIIO4 tetrahedra and the other PO4 tetrahedra, leading to the formation of a second layer also lying parallel to the (001) plane. In the first layer, all tetrahedra are oriented downwards, while in the second, they are oriented upwards with respect to the c axis. The interconnection between these two layers is ensured by the sharing of the vertices of the CoO4 and PO4 tetrahedra, as well as the vertex of the FeO4 tetrahedra, with those of the other PO4 tetrahedra. The stacking of these two layers leads to the formation of [CoFeP2O8]−∞ double layers lying parallel to the (001) plane and creates hexagonal tunnels running parallel to the c-axis direction. The Rb+ cations are located at the intersection of the tunnels with the interlayer space, thus forming a plane lying perpendicular to the c-axis direction, sandwiched between the double layers. In this structure, each rubidium atom exhibits a coordination number of twelve, whereas in the similar structure of KMgFe(PO4)2, which has monoclinic symmetry, the potassium cation is surrounded by eight oxygen atoms.
Keywords: phosphate; sheet; crystal structure; X-ray diffraction.
CCDC reference: 2576725
1. Chemical context
In solid-state chemistry and materials science, transition-metal (TM) phosphates have been studied extensively, particularly those with open structures. These porous materials are interesting due to their potential industrial and environmental applications, ranging from catalysis to ion exchange and separation (Gier & Stucky, 1991 ; Maspoch et al., 2007; Lin et al., 2004), electrochemistry (Trad et al., 2010) and magnetism (Ofer et al., 2012; Hadouchi et al., 2019). Among these, iron phosphates are particularly attractive due to their rich crystal chemistry (Moore, 1970; Redrup & Weller, 2009; Lajmi et al., 2009) and variable physical properties (Elbouaanani et al., 2002; Riou-Cavellec et al., 1999). The structural diversity is mainly due to the ability of TM cations to adopt different oxidation states such as PbMnII2MnIII(PO4)3 (Alhakmi et al., 2013) and coordination geometries, leading to a wide variety of families such as NaZnAl(PO4)2 (Yakubovich, et al., 2019) and NaCuIn(PO4)2 (Benhsina et al., 2020).
Our research team has previously developed and characterized several new phosphates with the general formula AMM′(PO4)2, where A, M, and M′ represent alkali or alkaline earth cations, divalent and trivalent metals, respectively. Most of the compounds were obtained by solid-state reactions and identified by X-ray diffraction. Investigations of these materials reveal the richness and diversity of their structures. The most frequent structural types adopted by these crystals are alluaudite, ABW-zeolite, α-CrPO4, and others. The following compounds represent an example of each structural type, respectively, Na2M2Fe(PO4)3 (M = Co, Ni) (Bouraima et al., 2015); β-KCoFe(PO4)2 (Bouraima et al., 2022); BaNi2Fe(PO4)3 (Ouaatta et al., 2023) and K3Ni6Fe(PO4)6 (Ouaatta et al., 2019). As part of our ongoing studies in this area, we now describe the synthesis and characterization of the title compound RbCoFe(PO4)2 (I) (Fig. 1).
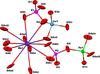 | Figure 1 The principal polyhedra constituting the crystal structure of the compound RbCoFe(PO4)2, with atomic labels. Displacement ellipsoids are represented at the 50% probability level. Symmetry codes:(i) −y + 2, x − y + 1, z; (ii) −x + y + 1, −x + 2, z; (iii) −y + 1, x − y, z; (iv) −x + y + 1, −x + 1, z; (v) x, y − 1, z − 1; (vi) x − 1, y, z; (vii) −y + 1, x − y, z; (viii) −y + 1, x − y + 1, z; (ix) −x + y, −x + 1, z; (x) x, y, z + 1; (xi) x, y + 1, z; (xii) −y, x − y, z; (xiii) −y + 1, x − y − 1, z; (xiv) −x + y + 2, −x + 1, z. |
2. Structural commentary
In this structure, all atoms occupy special positions of the P3m1 space group. The asymmetric unit of the new phase consists of one Rb atom in Wyckoff position 1b (3m), Co1, P2, and O3 atoms in position 1c (3m), Fe1, P1, and O1 atoms in position 1a (3m), while O2 and O4 occupy Wyckoff position 3d (m). However, refining the structure with cobalt in Wyckoff position (1a) and iron in (1c) leads to inconclusive results. Reversing the positions of Fe in (1a) and Co in (1c) leads to slightly better results: the Co—O distances (Table 1) are slightly longer than the Fe—O distances, and the bond-valence sums are very close to the expected values. It is also noted that freely refining the occupancies of the Co/Fe sites by a mixture of Co2+ and Fe3+ leads to unstable and the diverges. Refining the statistical distribution of the two cations Co2+ and Fe3+ at the two Wyckoff sites (1a) and (1c) converged well, with a slight improvement in the values of R (0.030) and Rw2 (0.077), but with very high standard deviations in the Co/Fe—O bond lengths. The same results are obtained when refining the structure by fixing different occupancy rates of the two sites by Co2+ and Fe3+. In light of the preceding analyses, we can conclude that the best model is the one corresponding to Fe3+ at (1a) and Co2+ at (1c).
|
 ; (ii)
 ; (iii)
 ; (iv)
 ; (v)
 .
Furthermore, all cations adopt a tetrahedral environment, except for the rubidium ion, which is surrounded by twelve oxygen atoms. The average phosphorus–oxygen bond lengths are 1.538 and 1.528 Å for P1 and P2, respectively. The average bond length for cobalt–oxygen is 1.887 Å, and 1.847 Å for the iron–oxygen distance. The is built by two distinct types of sheets lying parallel to the ab plane. The CoO4 tetrahedra alternate with P1O4 tetrahedra, sharing vertices and extending along both the a- and b-axis directions to construct a first tetrahedral sheet. All tetrahedra within this layer are oriented upward relative to the c axis. These sheets exhibit hexagonal-shaped pores. The same alternation is observed between FeO4, and P2O4 tetrahedra, leading to the formation of a second sheet, also encapsulating hexagonal-shaped pores. In this layer, the FeO4 and P2O4 tetrahedra are oriented ‘down' relative to the c axis. The interconnection between these two sheets is ensured through vertex-sharing between CoO4 and P2O4, and FeO4 and P1O4 tetrahedra via oxygen atoms (these inter-sheet bonds are constrained to be linear by symmetry). This arrangement leads to a [CoFeP2O8]−∞ double sheet lying parallel to the ab plane. The Rb+ cations form a plane sandwiched between the double layers. Each rubidium cation is surrounded by twelve oxygen atoms at the intersection of the tunnels with the interlayer space (Fig. 2).
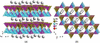 | Figure 2 (a) Perspective view of the crystal structure of RbCoFe(PO4)2, showing the plane of Rb+ ions sandwiched between the double layers of tetrahedra. (b) The Rb+ ions are located at the intersection of the channels extending along the [001] direction with the interlayer space. |
The reliability of the structural model was validated using bond-valence-sum (BVS; Altermatt & Brown, 1985) and charge-distribution (CHARDI; Nespolo, 2016) methods. The CHARDI and BVS computations were performed with the CHARDI2015 (Nespolo & Guillot, 2016) and EXPO2014 (Altomare et al., 2013) programs, respectively. The resulting parameters derived from both methods are listed in Table 2. The computed values V(i) (valence) and Q(i) (cationic charge) reveal excellent agreement between these values and their corresponding weighted oxidation numbers q(i)·sof(i). In accordance with this charge-distribution scheme, the cationic charge ratio q(i)/Q(i) ≃ 1 for all species, confirming the accuracy of this structural determination (Nespolo et al., 1999). This model is also corroborated by the low of the mean absolute percentage deviation (MAPD = 2.9%), which assesses the agreement between the calculated CHARDI charges Q(i) and the expected formal ones q(i) (Eon & Nespolo, 2015).
| |||||||||||||||||||||||||||||||||||||||||||||||||||
In the of RbCoFe(PO4)2, the coordination numbers (CN) and effective coordination numbers (ECoN) found for all the cationic sites are entirely consistent with the expected environments. For all crystallographic sites, the CN and ECoN values closely match, indicating well-defined coordination polyhedra, with the exception of the Rb1 atom, which exhibits a very low distortion level. Indeed, the Rb1 site adopts a high coordination number (CN = 12) with an ECoN value of 11.99, reflecting an almost ideal twelvefold coordination environment. This result confirms the existence of weakly directional bonds typical of alkali metal cations, where slight variations in bond lengths have a negligible impact on effective coordination. Both transition-metal sites, Co1 and Fe1, exhibit the same CN and ECoN values (CN = ECoN = 4.00), a feature of regular metal–oxygen bond lengths and consequently well-defined tetrahedral coordination geometries. Similarly, the P1 and P2 sites, each display CN = 4 and ECoN = 4.00, confirming the presence of highly regular PO4 tetrahedra.
Comparison of BVS results with the nominal valences V(i) confirms that the BVS approach leads to the expected values for the different crystallographic sites in the crystal structure of RbCoFe(PO4)2. The global instability index (GII) was also calculated to assess the electrostatic plausibility and the overall stability of the crystal structure (Salinas-Sanchez et al., 1992). The resulting GII value of 0.108 suggests a structurally stable and rigid framework, thereby confirming the reliability of the structural model.
The title compound crystallizes in a non-centrosymmetric space group with a polar axis (P3m1), which allows it to be classified among compounds likely to have ferroelectric properties (Abrahams, 2008). The ferroelectric paraelectric phase transition requires a space group change from P3m1 to one of its the non-isomorphic supergroups P m1, P
 m2, P63mc, or P6mm. Analysis of atomic positions using PLATON (Spek, 2020) shows that the structure could exhibit an approximate/pseudo-inversion symmetry (Marsh, 1994).
3. Database survey
A survey of the Powder Diffraction database (PDF2; version 2023) and of the Inorganic Database (ICSD; version 5.5.0 from FIZ Karlsruhe) shows that among phosphates of the type AMM′(PO4)2 (A = K or Rb, M = divalent cation and M′ = trivalent cation), to our knowledge only RbCoFe(PO4)2 crystallizes in a trigonal symmetry (space group P3m1) with a = 5.4474 Å and c = 8.1547 Å.
A number of known phosphates show similar unit cells, such as: β-CaZn2(PO4)2 (a = 5.180 Å, c = 7.670 Å, P3; Kreidler & Hummel, 1967; Czaya, 1972), SrMg2(PO4)2 (a = 5.133 Å, c = 8.038 Å, P3m; CPDS N 52–1587), and BaMg2(PO4)2 (a = 5.236 Å, c = 8.201 Å, P3m; JCPDS N 52–1587), space group P3m1), β-NaZnPO4 (a = 5.101 Å, c = 8.365 Å, P63; JCPDS N 52-0483) and NaBaPO4 (a = 5.622 Å, c = 7.259 Å, P3m1; Calvo & Faggiani, 1975; Launay et al., 1992). Despite these similarities, the structures of these compounds differ from that studied here. In these materials, the polyhedra form rings around the large cations, whereas in the structure of RbCoFe(PO4)2, the tetrahedra are ordered and form a double layer surrounding the rubidium plane. It may be noted that this lamellar structure is adopted by AMFe(PO4)2 phosphates (M = Mg, Co, Zn and A = K or Rb), although the latter crystallize in the monoclinic system, C2/c.
4. Synthesis and crystallization
The title compound was synthesized by mixing cobalt nitrate [Co(NO3)·6H2O], iron nitrate [Fe(NO3)3·9H2O], orthophosphoric acid (H3PO4) and rubidium carbonate (Rb2CO3) in the molar ratio 1:1:1:2. The mixture was placed in a small beaker containing distilled water, stirred for 24 h at room temperature, and subsequently evaporated to dryness. The resulting reaction mixture was heat-treated at 573 K and 773 K before being thoroughly ground in an agate mortar. The resulting solid was then transferred into a platinum crucible and heated to the melting temperature of 1233 K. After holding this temperature for 15 min, the molten product was cooled gradually to 1073 K at a rate of 5 K h−1, and then quickly to room temperature by shutting off the furnace. The final product comprised purple single crystals of suitable size for X-ray diffraction analysis, corresponding to the title phosphate.
5. Refinement
Crystal data, data collection and structure details are given in Table 3. The crystal structure was refined as an inversion twin using the TWIN/BASF instructions in SHELXL. The refined Flack parameter is 0.50 (7), indicating a near-perfect two-component inversion twin. The largest residual electron-density peak and the deepest hole in the last difference-Fourier map are located 0.53 and 0.17 Å from atoms O2 and O4, respectively.
|
Supporting information
CCDC reference: 2576725
contains datablock I. DOI: https://doi.org/10.1107/S2056989026007632/oo2018sup1.cif
Structure factors: contains datablock I. DOI: https://doi.org/10.1107/S2056989026007632/oo2018Isup2.hkl
| RbCoFe(PO4)2 | Dx = 3.092 Mg m−3 |
| Mr = 390.19 | Mo Kα radiation, λ = 0.71073 Å |
| Trigonal, P3m1 | Cell parameters from 945 reflections |
| a = 5.4474 (2) Å | θ = 2.5–37.8° |
| c = 8.1547 (3) Å | µ = 9.87 mm−1 |
| V = 209.56 (2) Å3 | T = 293 K |
| Z = 1 | Block, purple |
| F(000) = 184 | 0.25 × 0.21 × 0.14 mm |
| Bruker X8 APEX Diffractometer | 945 independent reflections |
| Radiation source: fine-focus sealed tube | 844 reflections with I > 2σ(I) |
| Graphite monochromator | Rint = 0.039 |
| φ and ω scans | θmax = 37.8°, θmin = 2.5° |
| Absorption correction: multi-scan (SADABS; Krause et al., 2015) | h = −9→9 |
| Tmin = 0.507, Tmax = 0.748 | k = −9→9 |
| 14543 measured reflections | l = −14→14 |
| Refinement on F2 | Primary atom site location: dual |
| Least-squares matrix: full | w = 1/[σ2(Fo2) + (0.0284P)2 + 0.4865P] where P = (Fo2 + 2Fc2)/3 |
| R[F2 > 2σ(F2)] = 0.032 | (Δ/σ)max < 0.001 |
| wR(F2) = 0.079 | Δρmax = 1.30 e Å−3 |
| S = 1.11 | Δρmin = −0.95 e Å−3 |
| 945 reflections | Absolute structure: Refined as an inversion twin |
| 35 parameters | Absolute structure parameter: 0.50 (7) |
| 4 restraints |
Geometry. All esds (except the esd in the dihedral angle between two l.s. planes) are estimated using the full covariance matrix. The cell esds are taken into account individually in the estimation of esds in distances, angles and torsion angles; correlations between esds in cell parameters are only used when they are defined by crystal symmetry. An approximate (isotropic) treatment of cell esds is used for estimating esds involving l.s. planes. |
Refinement. Refined as a 2-component inversion twin. |
| x | y | z | Uiso*/Ueq | ||
| Rb1 | 0.333333 | 0.666667 | 0.4375 (7) | 0.02836 (19) | |
| Fe1 | 1.000000 | 0.000000 | 0.1327 (6) | 0.0167 (6) | |
| Co1 | 0.666667 | 0.333333 | 0.7444 (6) | 0.0204 (6) | |
| P1 | 1.000000 | 1.000000 | 0.7177 (5) | 0.0198 (11) | |
| P2 | 0.666667 | 0.333333 | 0.1577 (4) | 0.0127 (8) | |
| O1 | 1.000000 | 1.000000 | 0.9066 (5) | 0.029 (5) | |
| O2 | 0.8489 (5) | 0.6978 (9) | 0.6473 (11) | 0.052 (4) | |
| O3 | 0.666667 | 0.333333 | −0.0263 (12) | 0.033 (6) | |
| O4 | 0.8199 (4) | 0.1801 (4) | 0.2218 (10) | 0.055 (4) |
| U11 | U22 | U33 | U12 | U13 | U23 | |
| Rb1 | 0.0310 (3) | 0.0310 (3) | 0.0230 (3) | 0.01552 (13) | 0.000 | 0.000 |
| Fe1 | 0.0175 (9) | 0.0175 (9) | 0.0153 (13) | 0.0087 (4) | 0.000 | 0.000 |
| Co1 | 0.0207 (9) | 0.0207 (9) | 0.0198 (14) | 0.0104 (4) | 0.000 | 0.000 |
| P1 | 0.0211 (15) | 0.0211 (15) | 0.017 (2) | 0.0106 (7) | 0.000 | 0.000 |
| P2 | 0.0121 (11) | 0.0121 (11) | 0.0139 (18) | 0.0061 (5) | 0.000 | 0.000 |
| O1 | 0.033 (7) | 0.033 (7) | 0.020 (7) | 0.016 (4) | 0.000 | 0.000 |
| O2 | 0.083 (9) | 0.018 (5) | 0.035 (5) | 0.009 (2) | 0.0034 (18) | 0.007 (4) |
| O3 | 0.044 (9) | 0.044 (9) | 0.011 (5) | 0.022 (4) | 0.000 | 0.000 |
| O4 | 0.099 (10) | 0.099 (10) | 0.019 (3) | 0.088 (10) | −0.0049 (17) | 0.0049 (17) |
| Rb1—O2i | 3.220 (5) | Fe1—O4xi | 1.848 (6) |
| Rb1—O2ii | 3.220 (5) | Fe1—O4 | 1.848 (6) |
| Rb1—O2iii | 3.220 (5) | Co1—O3xii | 1.870 (13) |
| Rb1—O2 | 3.220 (5) | Co1—O2vi | 1.893 (7) |
| Rb1—O2iv | 3.220 (5) | Co1—O2v | 1.893 (7) |
| Rb1—O2v | 3.220 (5) | Co1—O2 | 1.893 (7) |
| Rb1—O4vi | 3.245 (5) | P1—O2 | 1.5373 (14) |
| Rb1—O4v | 3.245 (5) | P1—O2xiii | 1.5373 (15) |
| Rb1—O4iii | 3.245 (5) | P1—O2i | 1.5373 (14) |
| Rb1—O4vii | 3.245 (5) | P1—O1 | 1.5398 (15) |
| Rb1—O4viii | 3.245 (5) | P2—O3 | 1.500 (9) |
| Rb1—O4i | 3.245 (5) | P2—O4 | 1.5374 (14) |
| Fe1—O1ix | 1.844 (9) | P2—O4v | 1.5374 (14) |
| Fe1—O4x | 1.848 (6) | P2—O4vi | 1.5374 (14) |
| O2i—Rb1—O2ii | 55.1 (2) | O4x—Fe1—Rb1xvi | 52.81 (12) |
| O2i—Rb1—O2iii | 94.4 (2) | O4xi—Fe1—Rb1xvi | 52.81 (12) |
| O2ii—Rb1—O2iii | 45.10 (12) | O4—Fe1—Rb1xvi | 118.5 (2) |
| O2i—Rb1—O2 | 45.10 (12) | Rb1xiv—Fe1—Rb1xvi | 85.60 (7) |
| O2ii—Rb1—O2 | 94.4 (2) | Rb1xv—Fe1—Rb1xvi | 85.60 (7) |
| O2iii—Rb1—O2 | 115.6 (3) | O3xii—Co1—O2vi | 114.7 (2) |
| O2i—Rb1—O2iv | 115.6 (3) | O3xii—Co1—O2v | 114.7 (2) |
| O2ii—Rb1—O2iv | 94.4 (2) | O2vi—Co1—O2v | 103.7 (3) |
| O2iii—Rb1—O2iv | 55.1 (2) | O3xii—Co1—O2 | 114.7 (2) |
| O2—Rb1—O2iv | 94.4 (2) | O2vi—Co1—O2 | 103.7 (3) |
| O2i—Rb1—O2v | 94.4 (2) | O2v—Co1—O2 | 103.7 (3) |
| O2ii—Rb1—O2v | 115.6 (3) | O3xii—Co1—Rb1 | 128.51 (5) |
| O2iii—Rb1—O2v | 94.4 (2) | O2vi—Co1—Rb1 | 116.8 (2) |
| O2—Rb1—O2v | 55.1 (2) | O2v—Co1—Rb1 | 51.99 (12) |
| O2iv—Rb1—O2v | 45.10 (12) | O2—Co1—Rb1 | 51.99 (12) |
| O2i—Rb1—O4vi | 125.48 (7) | O3xii—Co1—Rb1xv | 128.51 (5) |
| O2ii—Rb1—O4vi | 179.2 (3) | O2vi—Co1—Rb1xv | 51.99 (12) |
| O2iii—Rb1—O4vi | 134.74 (4) | O2v—Co1—Rb1xv | 51.99 (12) |
| O2—Rb1—O4vi | 86.37 (11) | O2—Co1—Rb1xv | 116.8 (2) |
| O2iv—Rb1—O4vi | 85.85 (12) | Rb1—Co1—Rb1xv | 85.33 (7) |
| O2v—Rb1—O4vi | 65.15 (8) | O3xii—Co1—Rb1xiv | 128.51 (5) |
| O2i—Rb1—O4v | 85.85 (12) | O2vi—Co1—Rb1xiv | 51.99 (12) |
| O2ii—Rb1—O4v | 134.74 (4) | O2v—Co1—Rb1xiv | 116.8 (2) |
| O2iii—Rb1—O4v | 179.2 (3) | O2—Co1—Rb1xiv | 51.99 (12) |
| O2—Rb1—O4v | 65.15 (8) | Rb1—Co1—Rb1xiv | 85.33 (7) |
| O2iv—Rb1—O4v | 125.48 (7) | Rb1xv—Co1—Rb1xiv | 85.33 (7) |
| O2v—Rb1—O4v | 86.37 (11) | O2—P1—O2xiii | 106.9 (5) |
| O4vi—Rb1—O4v | 45.39 (10) | O2—P1—O2i | 106.9 (5) |
| O2i—Rb1—O4iii | 179.2 (3) | O2xiii—P1—O2i | 106.9 (5) |
| O2ii—Rb1—O4iii | 125.48 (7) | O2—P1—O1 | 111.9 (4) |
| O2iii—Rb1—O4iii | 86.37 (11) | O2xiii—P1—O1 | 111.9 (4) |
| O2—Rb1—O4iii | 134.74 (4) | O2i—P1—O1 | 111.9 (4) |
| O2iv—Rb1—O4iii | 65.15 (8) | O2—P1—Rb1 | 53.5 (2) |
| O2v—Rb1—O4iii | 85.85 (12) | O2xiii—P1—Rb1 | 122.1 (5) |
| O4vi—Rb1—O4iii | 53.9 (2) | O2i—P1—Rb1 | 53.5 (2) |
| O4v—Rb1—O4iii | 93.4 (2) | O1—P1—Rb1 | 126.00 (11) |
| O2i—Rb1—O4vii | 65.15 (8) | O2—P1—Rb1xvii | 122.1 (5) |
| O2ii—Rb1—O4vii | 86.37 (11) | O2xiii—P1—Rb1xvii | 53.5 (2) |
| O2iii—Rb1—O4vii | 125.48 (7) | O2i—P1—Rb1xvii | 53.5 (2) |
| O2—Rb1—O4vii | 85.85 (12) | O1—P1—Rb1xvii | 126.00 (11) |
| O2iv—Rb1—O4vii | 179.2 (3) | Rb1—P1—Rb1xvii | 88.95 (15) |
| O2v—Rb1—O4vii | 134.74 (4) | O2—P1—Rb1xiv | 53.5 (2) |
| O4vi—Rb1—O4vii | 93.4 (2) | O2xiii—P1—Rb1xiv | 53.5 (2) |
| O4v—Rb1—O4vii | 53.9 (2) | O2i—P1—Rb1xiv | 122.1 (5) |
| O4iii—Rb1—O4vii | 114.1 (3) | O1—P1—Rb1xiv | 126.00 (11) |
| O2i—Rb1—O4viii | 134.74 (4) | Rb1—P1—Rb1xiv | 88.95 (15) |
| O2ii—Rb1—O4viii | 85.85 (12) | Rb1xvii—P1—Rb1xiv | 88.95 (15) |
| O2iii—Rb1—O4viii | 65.15 (8) | O3—P2—O4 | 109.9 (4) |
| O2—Rb1—O4viii | 179.2 (3) | O3—P2—O4v | 109.9 (4) |
| O2iv—Rb1—O4viii | 86.37 (11) | O4—P2—O4v | 109.1 (4) |
| O2v—Rb1—O4viii | 125.48 (7) | O3—P2—O4vi | 109.9 (4) |
| O4vi—Rb1—O4viii | 93.4 (2) | O4—P2—O4vi | 109.1 (4) |
| O4v—Rb1—O4viii | 114.1 (3) | O4v—P2—O4vi | 109.1 (4) |
| O4iii—Rb1—O4viii | 45.39 (10) | O3—P2—Rb1xiv | 125.96 (9) |
| O4vii—Rb1—O4viii | 93.4 (2) | O4—P2—Rb1xiv | 54.5 (2) |
| O2i—Rb1—O4i | 86.37 (11) | O4v—P2—Rb1xiv | 54.5 (2) |
| O2ii—Rb1—O4i | 65.15 (8) | O4vi—P2—Rb1xiv | 124.2 (4) |
| O2iii—Rb1—O4i | 85.85 (12) | O3—P2—Rb1xv | 125.96 (9) |
| O2—Rb1—O4i | 125.48 (7) | O4—P2—Rb1xv | 54.5 (2) |
| O2iv—Rb1—O4i | 134.74 (4) | O4v—P2—Rb1xv | 124.2 (4) |
| O2v—Rb1—O4i | 179.2 (3) | O4vi—P2—Rb1xv | 54.5 (2) |
| O4vi—Rb1—O4i | 114.1 (3) | Rb1xiv—P2—Rb1xv | 89.01 (13) |
| O4v—Rb1—O4i | 93.4 (2) | O3—P2—Rb1 | 125.96 (9) |
| O4iii—Rb1—O4i | 93.4 (2) | O4—P2—Rb1 | 124.2 (4) |
| O4vii—Rb1—O4i | 45.39 (10) | O4v—P2—Rb1 | 54.5 (2) |
| O4viii—Rb1—O4i | 53.9 (2) | O4vi—P2—Rb1 | 54.5 (2) |
| O1ix—Fe1—O4x | 113.1 (2) | Rb1xiv—P2—Rb1 | 89.01 (13) |
| O1ix—Fe1—O4xi | 113.1 (2) | Rb1xv—P2—Rb1 | 89.01 (13) |
| O4x—Fe1—O4xi | 105.6 (3) | P1—O1—Fe1xviii | 180.0 |
| O1ix—Fe1—O4 | 113.1 (2) | P1—O2—Co1 | 133.3 (6) |
| O4x—Fe1—O4 | 105.6 (3) | P1—O2—Rb1 | 103.9 (3) |
| O4xi—Fe1—O4 | 105.6 (3) | Co1—O2—Rb1 | 100.42 (9) |
| O1ix—Fe1—Rb1xiv | 128.32 (5) | P1—O2—Rb1xiv | 103.9 (3) |
| O4x—Fe1—Rb1xiv | 52.81 (12) | Co1—O2—Rb1xiv | 100.42 (9) |
| O4xi—Fe1—Rb1xiv | 118.5 (2) | Rb1—O2—Rb1xiv | 115.6 (3) |
| O4—Fe1—Rb1xiv | 52.81 (12) | P2—O3—Co1xix | 180.0 |
| O1ix—Fe1—Rb1xv | 128.32 (5) | P2—O4—Fe1 | 137.0 (6) |
| O4x—Fe1—Rb1xv | 118.5 (2) | P2—O4—Rb1xiv | 102.8 (2) |
| O4xi—Fe1—Rb1xv | 52.81 (12) | Fe1—O4—Rb1xiv | 100.21 (8) |
| O4—Fe1—Rb1xv | 52.81 (12) | P2—O4—Rb1xv | 102.8 (2) |
| Rb1xiv—Fe1—Rb1xv | 85.60 (7) | Fe1—O4—Rb1xv | 100.21 (8) |
| O1ix—Fe1—Rb1xvi | 128.32 (5) | Rb1xiv—O4—Rb1xv | 114.1 (3) |
| Symmetry codes: (i) −x+y+1, −x+2, z; (ii) −y+1, x−y+1, z; (iii) x−1, y, z; (iv) −x+y, −x+1, z; (v) −y+1, x−y, z; (vi) −x+y+1, −x+1, z; (vii) x, y+1, z; (viii) −y, x−y, z; (ix) x, y−1, z−1; (x) −x+y+2, −x+1, z; (xi) −y+1, x−y−1, z; (xii) x, y, z+1; (xiii) −y+2, x−y+1, z; (xiv) x+1, y, z; (xv) x, y−1, z; (xvi) x+1, y−1, z; (xvii) x+1, y+1, z; (xviii) x, y+1, z+1; (xix) x, y, z−1. |
| q(i) = formal oxidation number; sof(i) = site occupancy; CN(i) = classical coordination number; Q(i) = calculated charge; V(i) = calculated valence; ECoN(i) = effective coordination number. |
| Cation | q(i)·sof(i) | CN(i) | ECoN(i) | V(i) | Q(i) | q(i)/Q(i) |
| Rb1 | 1 | 12 | 11.99 | 0.87 | 1.00 | 1.00 |
| Co1 | 2 | 4 | 4.00 | 2.02 | 2.00 | 0.96 |
| Fe1 | 3 | 4 | 4.00 | 3.17 | 2.88 | 1.04 |
| P1 | 5 | 4 | 4.00 | 4.95 | 5.17 | 0.97 |
| P2 | 5 | 4 | 3.98 | 5.09 | 4.87 | 1.03 |
Acknowledgements
The authors would like to express their sincere gratitude to the Technical and Scientific Research Support Unit (UATRS, CNRST) for the X-ray measurements and to Mohammed V University in Rabat, Morocco, for the financial support.
References
 Abrahams, S. C. (2008). Acta Cryst. B64, 426–437. Web of Science CrossRef CAS IUCr Journals Google Scholar
Alhakmi, G., Assani, A., Saadi, M. & El Ammari, L. (2013). Acta Cryst. E69, i40. CrossRef IUCr Journals Google Scholar
Altermatt, D. & Brown, I. D. (1985). Acta Cryst. B41, 240–244. CrossRef CAS Web of Science IUCr Journals Google Scholar
Altomare, A., Cuocci, C., Giacovazzo, C., Moliterni, A., Rizzi, R., Corriero, N. & Falcicchio, A. (2013). J. Appl. Cryst. 46, 1231–1235. Web of Science CrossRef CAS IUCr Journals Google Scholar
Benhsina, E., Khmiyas, J., Ouaatta, S., Assani, A., Saadi, M. & El Ammari, L. (2020). Acta Cryst. E76, 366–369. Web of Science CrossRef ICSD IUCr Journals Google Scholar
Bouraima, A., Assani, A., Saadi, M., Makani, T. & El Ammari, L. (2015). Acta Cryst. E71, 558–560. Web of Science CrossRef ICSD IUCr Journals Google Scholar
Bouraima, A., Ouaatta, S., Khmiyas, J., Anguilè, J. J., Makani, T., Assani, A., Saadi, M. & El Ammari, L. (2022). Acta Cryst. E78, 746–749. CrossRef IUCr Journals Google Scholar
Brandenburg, K. (2006). DIAMOND. Crystal Impact GbR, Bonn, Germany. Google Scholar
Bruker, (2016). APEX3 and SAINT. Bruker AXS Inc., Madison, Wisconsin, USA. Google Scholar
Calvo, C. & Faggiani, R. (1975). Can. J. Chem. 53, 1849–1853. CrossRef Google Scholar
Czaya, R. (1972). Acta Cryst. B28, 322–323. CrossRef IUCr Journals Google Scholar
Elbouaanani, L. K., Malaman, B., Gérardin, R. & Ijjaali, M. (2002). J. Solid State Chem. 163, 412–420. Web of Science CrossRef CAS Google Scholar
Eon, J.-G. & Nespolo, M. (2015). Acta Cryst. B71, 34–47. Web of Science CrossRef IUCr Journals Google Scholar
Farrugia, L. J. (2012). J. Appl. Cryst. 45, 849–854. Web of Science CrossRef CAS IUCr Journals Google Scholar
Gier, T. E. & Stucky, G. D. (1991). Nature 349, 508–510. CrossRef Google Scholar
Hadouchi, M., Assani, A., Saadi, M., Lahmar, A., El Marssi, M., Sajieddine, M. & El Ammari, L. (2019). J. Supercond. Nov. Magn. 32, 2437–2446. CrossRef Google Scholar
Krause, L., Herbst-Irmer, R., Sheldrick, G. M. & Stalke, D. (2015). J. Appl. Cryst. 48, 3–10. Web of Science CSD CrossRef ICSD CAS IUCr Journals Google Scholar
Kreidler, E. R. & Hummel, F. A. (1967). Inorg. Chem. 6, 524–528. CrossRef Google Scholar
Lajmi, B., Hidouri, M., Hammouda, A. B., Wattiaux, A., Fournés, L., Darriet, J. & Amara, M. B. (2009). Mater. Chem. Phys. 113, 372–375. CrossRef Google Scholar
Launay, S., Mahé, P. & Quarton, M. (1992). Mater. Res. Bull. 27, 1347–1353. CrossRef Google Scholar
Lin, D.-C., Xu, X.-W., Zuo, F. & Long, Y.-C. (2004). Microporous Mesoporous Mater. 70, 63–70. CrossRef Google Scholar
Marsh, R. E. (1994). Acta Cryst. C50, 1713–1715. CrossRef IUCr Journals Google Scholar
Maspoch, D., Ruiz-Molina, D. & Veciana, J. (2007). Chem. Soc. Rev. 36, 770–818. Web of Science CrossRef PubMed CAS Google Scholar
Moore, P. B. (1970). Am. Mineral. 55, 135–169. CAS Google Scholar
Nespolo, M. (2016). Acta Cryst. B72, 51–66. Web of Science CrossRef IUCr Journals Google Scholar
Nespolo, M., Ferraris, G. & Ohashi, H. (1999). Acta Cryst. B55, 902–916. Web of Science CrossRef CAS IUCr Journals Google Scholar
Nespolo, M. & Guillot, B. (2016). J. Appl. Cryst. 49, 317–321. Web of Science CrossRef CAS IUCr Journals Google Scholar
Ofer, O., Sugiyama, J., Brewer, J. H., Månsson, M., Prša, K., Ansaldo, E. J., Kobayashi, G. & Kanno, R. (2012). Phys. Procedia 30, 160–163. CrossRef Google Scholar
Ouaatta, S., Assani, A., Saadi, M. & El Ammari, L. (2019). Acta Cryst. E75, 402–404. Web of Science CrossRef ICSD IUCr Journals Google Scholar
Ouaatta, S., Bouraima, A., Benhsina, E., Khmiyas, J., Assani, A., Saadi, M. & El Ammari, L. (2023). Acta Cryst. E79, 95–98. CrossRef IUCr Journals Google Scholar
Redrup, K. V. & Weller, M. T. (2009). Dalton Trans. pp. 3786–3792. Web of Science CrossRef Google Scholar
Riou-Cavellec, M., Riou, D. & Férey, G. (1999). Inorg. Chim. Acta 291, 317–325. Google Scholar
Salinas-Sanchez, A., Garcia-Muñoz, J. L., Rodriguez-Carvajal, J., Saez-Puche, R. & Martinez, J. L. (1992). J. Solid State Chem. 100, 201–211. CAS Google Scholar
Sheldrick, G. M. (2015a). Acta Cryst. A71, 3–8. Web of Science CrossRef IUCr Journals Google Scholar
Sheldrick, G. M. (2015b). Acta Cryst. C71, 3–8. Web of Science CrossRef IUCr Journals Google Scholar
Spek, A. L. (2020). Acta Cryst. E76, 1–11. Web of Science CrossRef IUCr Journals Google Scholar
Trad, K., Carlier, D., Croguennec, L., Wattiaux, A., Ben Amara, M. & Delmas, C. (2010). Chem. Mater. 22, 5554–5562. Web of Science CrossRef CAS Google Scholar
Westrip, S. P. (2010). J. Appl. Cryst. 43, 920–925. Web of Science CrossRef CAS IUCr Journals Google Scholar
Yakubovich, O., Kiriukhina, G., Volkov, A. & Dimitrova, O. (2019). Acta Cryst. C75, 514–522. Web of Science CrossRef ICSD IUCr Journals Google Scholar
This is an open-access article distributed under the terms of the Creative Commons Attribution (CC-BY) Licence, which permits unrestricted use, distribution, and reproduction in any medium, provided the original authors and source are cited.
| CRYSTALLOGRAPHIC COMMUNICATIONS |